Jump to
S1C2
S1C3
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -210.062170 |
| Energy at 298.15K | -210.071395 |
| HF Energy | -209.189671 |
| Nuclear repulsion energy | 135.609045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3764 |
3573 |
75.21 |
|
|
|
| 2 |
A |
3651 |
3467 |
7.06 |
|
|
|
| 3 |
A |
3556 |
3376 |
1.70 |
|
|
|
| 4 |
A |
3149 |
2990 |
32.86 |
|
|
|
| 5 |
A |
3140 |
2981 |
23.33 |
|
|
|
| 6 |
A |
3063 |
2908 |
59.04 |
|
|
|
| 7 |
A |
3031 |
2878 |
58.19 |
|
|
|
| 8 |
A |
1659 |
1575 |
28.65 |
|
|
|
| 9 |
A |
1542 |
1464 |
0.80 |
|
|
|
| 10 |
A |
1523 |
1446 |
5.32 |
|
|
|
| 11 |
A |
1460 |
1386 |
61.74 |
|
|
|
| 12 |
A |
1419 |
1347 |
11.59 |
|
|
|
| 13 |
A |
1394 |
1324 |
6.91 |
|
|
|
| 14 |
A |
1345 |
1277 |
2.01 |
|
|
|
| 15 |
A |
1277 |
1212 |
26.02 |
|
|
|
| 16 |
A |
1208 |
1147 |
5.18 |
|
|
|
| 17 |
A |
1142 |
1084 |
48.73 |
|
|
|
| 18 |
A |
1092 |
1036 |
43.75 |
|
|
|
| 19 |
A |
1023 |
971 |
7.93 |
|
|
|
| 20 |
A |
947 |
899 |
47.53 |
|
|
|
| 21 |
A |
909 |
863 |
13.46 |
|
|
|
| 22 |
A |
833 |
791 |
87.95 |
|
|
|
| 23 |
A |
584 |
555 |
112.83 |
|
|
|
| 24 |
A |
542 |
515 |
12.09 |
|
|
|
| 25 |
A |
333 |
316 |
0.61 |
|
|
|
| 26 |
A |
269 |
256 |
12.98 |
|
|
|
| 27 |
A |
197 |
187 |
6.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22024.7 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20910.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.341 |
-0.561 |
0.118 |
| C2 |
-0.628 |
0.651 |
-0.274 |
| C3 |
0.779 |
0.555 |
0.271 |
| O4 |
1.394 |
-0.634 |
-0.168 |
| H5 |
-1.612 |
-0.510 |
1.090 |
| H6 |
-2.189 |
-0.674 |
-0.416 |
| H7 |
-1.098 |
1.577 |
0.061 |
| H8 |
-0.576 |
0.670 |
-1.357 |
| H9 |
1.378 |
1.389 |
-0.078 |
| H10 |
0.745 |
0.600 |
1.362 |
| H11 |
0.685 |
-1.289 |
-0.122 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4604 | 2.4005 | 2.7506 | 1.0103 | 1.0082 | 2.1524 | 2.0684 | 3.3516 | 2.6927 | 2.1658 |
C2 | 1.4604 | | 1.5113 | 2.3979 | 2.0434 | 2.0526 | 1.0903 | 1.0851 | 2.1458 | 2.1367 | 2.3476 | C3 | 2.4005 | 1.5113 | | 1.4084 | 2.7421 | 3.2843 | 2.1471 | 2.1207 | 1.0845 | 1.0933 | 1.8876 | O4 | 2.7506 | 2.3979 | 1.4084 | | 3.2607 | 3.5910 | 3.3387 | 2.6441 | 2.0247 | 2.0703 | 0.9665 | H5 | 1.0103 | 2.0434 | 2.7421 | 3.2607 | | 1.6211 | 2.3821 | 2.9076 | 3.7291 | 2.6196 | 2.7113 | H6 | 1.0082 | 2.0526 | 3.2843 | 3.5910 | 1.6211 | | 2.5461 | 2.3008 | 4.1338 | 3.6600 | 2.9529 | H7 | 2.1524 | 1.0903 | 2.1471 | 3.3387 | 2.3821 | 2.5461 | | 1.7626 | 2.4868 | 2.4587 | 3.3798 | H8 | 2.0684 | 1.0851 | 2.1207 | 2.6441 | 2.9076 | 2.3008 | 1.7626 | | 2.4430 | 3.0244 | 2.6365 | H9 | 3.3516 | 2.1458 | 1.0845 | 2.0247 | 3.7291 | 4.1338 | 2.4868 | 2.4430 | | 1.7601 | 2.7664 | H10 | 2.6927 | 2.1367 | 1.0933 | 2.0703 | 2.6196 | 3.6600 | 2.4587 | 3.0244 | 1.7601 | | 2.4035 | H11 | 2.1658 | 2.3476 | 1.8876 | 0.9665 | 2.7113 | 2.9529 | 3.3798 | 2.6365 | 2.7664 | 2.4035 | |
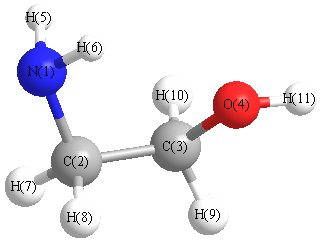 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.747 |
|
N1 |
C2 |
H7 |
114.316 |
| N1 |
C2 |
H8 |
107.785 |
|
C2 |
N1 |
H5 |
110.263 |
| C2 |
N1 |
H6 |
111.182 |
|
C2 |
C3 |
O4 |
110.372 |
| C2 |
C3 |
H9 |
110.430 |
|
C2 |
C3 |
H10 |
109.185 |
| C3 |
C2 |
H7 |
110.181 |
|
C3 |
C2 |
H8 |
108.411 |
| C3 |
O4 |
H11 |
103.708 |
|
O4 |
C3 |
H9 |
107.913 |
| O4 |
C3 |
H10 |
111.065 |
|
H5 |
N1 |
H6 |
106.864 |
| H7 |
C2 |
H8 |
108.237 |
|
H9 |
C3 |
H10 |
107.839 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
