Jump to
S1C2
S1C3
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -210.419716 |
| Energy at 298.15K | -210.428872 |
| HF Energy | -210.419716 |
| Nuclear repulsion energy | 135.261254 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3755 |
3602 |
73.68 |
|
|
|
| 2 |
A |
3635 |
3487 |
4.39 |
|
|
|
| 3 |
A |
3547 |
3403 |
0.57 |
|
|
|
| 4 |
A |
3098 |
2971 |
39.65 |
|
|
|
| 5 |
A |
3087 |
2961 |
31.62 |
|
|
|
| 6 |
A |
3006 |
2883 |
72.36 |
|
|
|
| 7 |
A |
2968 |
2846 |
68.94 |
|
|
|
| 8 |
A |
1659 |
1592 |
28.74 |
|
|
|
| 9 |
A |
1515 |
1454 |
0.59 |
|
|
|
| 10 |
A |
1497 |
1436 |
6.67 |
|
|
|
| 11 |
A |
1450 |
1391 |
72.36 |
|
|
|
| 12 |
A |
1409 |
1351 |
10.31 |
|
|
|
| 13 |
A |
1375 |
1319 |
6.98 |
|
|
|
| 14 |
A |
1332 |
1277 |
2.52 |
|
|
|
| 15 |
A |
1259 |
1208 |
26.26 |
|
|
|
| 16 |
A |
1198 |
1149 |
4.55 |
|
|
|
| 17 |
A |
1131 |
1085 |
65.12 |
|
|
|
| 18 |
A |
1085 |
1041 |
38.99 |
|
|
|
| 19 |
A |
1009 |
968 |
7.45 |
|
|
|
| 20 |
A |
932 |
894 |
49.57 |
|
|
|
| 21 |
A |
893 |
857 |
8.64 |
|
|
|
| 22 |
A |
819 |
786 |
91.17 |
|
|
|
| 23 |
A |
580 |
556 |
108.41 |
|
|
|
| 24 |
A |
536 |
514 |
14.19 |
|
|
|
| 25 |
A |
323 |
310 |
0.71 |
|
|
|
| 26 |
A |
266 |
255 |
11.35 |
|
|
|
| 27 |
A |
181 |
174 |
6.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21771.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 20883.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.349 |
-0.558 |
0.118 |
| C2 |
-0.630 |
0.654 |
-0.264 |
| C3 |
0.791 |
0.554 |
0.260 |
| O4 |
1.393 |
-0.641 |
-0.164 |
| H5 |
-1.642 |
-0.515 |
1.085 |
| H6 |
-2.179 |
-0.688 |
-0.440 |
| H7 |
-1.096 |
1.583 |
0.087 |
| H8 |
-0.597 |
0.691 |
-1.354 |
| H9 |
1.391 |
1.387 |
-0.110 |
| H10 |
0.780 |
0.622 |
1.359 |
| H11 |
0.685 |
-1.296 |
-0.119 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4601 | 2.4159 | 2.7579 | 1.0108 | 1.0087 | 2.1563 | 2.0720 | 3.3676 | 2.7322 | 2.1774 |
C2 | 1.4601 | | 1.5181 | 2.4044 | 2.0521 | 2.0569 | 1.0969 | 1.0911 | 2.1553 | 2.1505 | 2.3573 | C3 | 2.4159 | 1.5181 | | 1.4040 | 2.7822 | 3.2946 | 2.1563 | 2.1334 | 1.0911 | 1.1005 | 1.8919 | O4 | 2.7579 | 2.4044 | 1.4040 | | 3.2840 | 3.5831 | 3.3475 | 2.6740 | 2.0285 | 2.0710 | 0.9656 | H5 | 1.0108 | 2.0521 | 2.7822 | 3.2840 | | 1.6262 | 2.3875 | 2.9147 | 3.7739 | 2.6896 | 2.7336 | H6 | 1.0087 | 2.0569 | 3.2946 | 3.5831 | 1.6262 | | 2.5707 | 2.2890 | 4.1422 | 3.7026 | 2.9461 | H7 | 2.1563 | 1.0969 | 2.1563 | 3.3475 | 2.3875 | 2.5707 | | 1.7664 | 2.5023 | 2.4625 | 3.3924 | H8 | 2.0720 | 1.0911 | 2.1334 | 2.6740 | 2.9147 | 2.2890 | 1.7664 | | 2.4460 | 3.0432 | 2.6685 | H9 | 3.3676 | 2.1553 | 1.0911 | 2.0285 | 3.7739 | 4.1422 | 2.5023 | 2.4460 | | 1.7650 | 2.7744 | H10 | 2.7322 | 2.1505 | 1.1005 | 2.0710 | 2.6896 | 3.7026 | 2.4625 | 3.0432 | 1.7650 | | 2.4230 | H11 | 2.1774 | 2.3573 | 1.8919 | 0.9656 | 2.7336 | 2.9461 | 3.3924 | 2.6685 | 2.7744 | 2.4230 | |
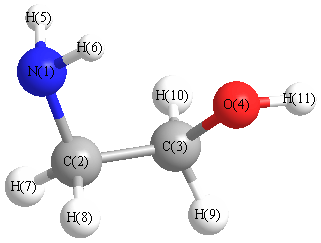 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.405 |
|
N1 |
C2 |
H7 |
114.229 |
| N1 |
C2 |
H8 |
107.740 |
|
C2 |
N1 |
H5 |
110.991 |
| C2 |
N1 |
H6 |
111.537 |
|
C2 |
C3 |
O4 |
110.677 |
| C2 |
C3 |
H9 |
110.316 |
|
C2 |
C3 |
H10 |
109.379 |
| C3 |
C2 |
H7 |
110.050 |
|
C3 |
C2 |
H8 |
108.589 |
| C3 |
O4 |
H11 |
104.433 |
|
O4 |
C3 |
H9 |
108.131 |
| O4 |
C3 |
H10 |
110.989 |
|
H5 |
N1 |
H6 |
107.264 |
| H7 |
C2 |
H8 |
107.677 |
|
H9 |
C3 |
H10 |
107.287 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.324 |
|
|
|
| 2 |
C |
-0.090 |
|
|
|
| 3 |
C |
0.078 |
|
|
|
| 4 |
O |
-0.361 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.059 |
|
|
|
| 8 |
H |
0.085 |
|
|
|
| 9 |
H |
0.063 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.200 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.834 |
1.022 |
0.523 |
3.057 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.166 |
2.102 |
0.312 |
| y |
2.102 |
-25.413 |
-0.270 |
| z |
0.312 |
-0.270 |
-24.416 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.252 |
2.102 |
0.312 |
| y |
2.102 |
-0.122 |
-0.270 |
| z |
0.312 |
-0.270 |
1.374 |
|
| Polar |
|---|
| 3z2-r2 | 2.748 |
|---|
| x2-y2 | -0.753 |
|---|
| xy | 2.102 |
|---|
| xz | 0.312 |
|---|
| yz | -0.270 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.037 |
0.185 |
0.034 |
| y |
0.185 |
5.703 |
0.050 |
| z |
0.034 |
0.050 |
5.204 |
<r2> (average value of r
2) Å
2
| <r2> |
86.049 |
| (<r2>)1/2 |
9.276 |
