Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -210.368113 |
| Energy at 298.15K | -210.377267 |
| HF Energy | -210.368113 |
| Nuclear repulsion energy | 135.527386 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3756 |
3594 |
75.89 |
|
|
|
| 2 |
A |
3635 |
3479 |
4.42 |
|
|
|
| 3 |
A |
3546 |
3394 |
0.60 |
|
|
|
| 4 |
A |
3103 |
2970 |
40.02 |
|
|
|
| 5 |
A |
3093 |
2960 |
32.75 |
|
|
|
| 6 |
A |
3011 |
2882 |
73.14 |
|
|
|
| 7 |
A |
2972 |
2844 |
70.51 |
|
|
|
| 8 |
A |
1655 |
1584 |
28.72 |
|
|
|
| 9 |
A |
1514 |
1450 |
0.55 |
|
|
|
| 10 |
A |
1495 |
1431 |
6.14 |
|
|
|
| 11 |
A |
1445 |
1383 |
69.89 |
|
|
|
| 12 |
A |
1405 |
1345 |
11.37 |
|
|
|
| 13 |
A |
1371 |
1313 |
6.89 |
|
|
|
| 14 |
A |
1328 |
1271 |
2.51 |
|
|
|
| 15 |
A |
1256 |
1202 |
26.51 |
|
|
|
| 16 |
A |
1194 |
1143 |
4.13 |
|
|
|
| 17 |
A |
1131 |
1082 |
68.18 |
|
|
|
| 18 |
A |
1085 |
1038 |
39.11 |
|
|
|
| 19 |
A |
1006 |
963 |
7.26 |
|
|
|
| 20 |
A |
930 |
890 |
48.81 |
|
|
|
| 21 |
A |
894 |
856 |
7.65 |
|
|
|
| 22 |
A |
817 |
782 |
91.38 |
|
|
|
| 23 |
A |
577 |
552 |
107.91 |
|
|
|
| 24 |
A |
529 |
506 |
14.39 |
|
|
|
| 25 |
A |
322 |
308 |
0.85 |
|
|
|
| 26 |
A |
264 |
253 |
10.88 |
|
|
|
| 27 |
A |
189 |
181 |
6.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21759.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20826.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.345 |
-0.556 |
0.119 |
| C2 |
-0.628 |
0.654 |
-0.266 |
| C3 |
0.789 |
0.553 |
0.262 |
| O4 |
1.387 |
-0.641 |
-0.165 |
| H5 |
-1.633 |
-0.512 |
1.086 |
| H6 |
-2.176 |
-0.687 |
-0.435 |
| H7 |
-1.094 |
1.582 |
0.079 |
| H8 |
-0.590 |
0.685 |
-1.354 |
| H9 |
1.390 |
1.384 |
-0.104 |
| H10 |
0.775 |
0.617 |
1.359 |
| H11 |
0.678 |
-1.293 |
-0.122 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4584 | 2.4092 | 2.7482 | 1.0097 | 1.0076 | 2.1536 | 2.0689 | 3.3608 | 2.7209 | 2.1662 |
C2 | 1.4584 | | 1.5155 | 2.3977 | 2.0491 | 2.0550 | 1.0947 | 1.0890 | 2.1523 | 2.1465 | 2.3491 | C3 | 2.4092 | 1.5155 | | 1.4024 | 2.7717 | 3.2886 | 2.1539 | 2.1286 | 1.0889 | 1.0984 | 1.8894 | O4 | 2.7482 | 2.3977 | 1.4024 | | 3.2722 | 3.5737 | 3.3409 | 2.6613 | 2.0259 | 2.0685 | 0.9650 | H5 | 1.0097 | 2.0491 | 2.7717 | 3.2722 | | 1.6244 | 2.3858 | 2.9111 | 3.7623 | 2.6736 | 2.7221 | H6 | 1.0076 | 2.0550 | 3.2886 | 3.5737 | 1.6244 | | 2.5662 | 2.2893 | 4.1372 | 3.6909 | 2.9340 | H7 | 2.1536 | 1.0947 | 2.1539 | 3.3409 | 2.3858 | 2.5662 | | 1.7642 | 2.4992 | 2.4622 | 3.3839 | H8 | 2.0689 | 1.0890 | 2.1286 | 2.6613 | 2.9111 | 2.2893 | 1.7642 | | 2.4440 | 3.0372 | 2.6534 | H9 | 3.3608 | 2.1523 | 1.0889 | 2.0259 | 3.7623 | 4.1372 | 2.4992 | 2.4440 | | 1.7626 | 2.7707 | H10 | 2.7209 | 2.1465 | 1.0984 | 2.0685 | 2.6736 | 3.6909 | 2.4622 | 3.0372 | 1.7626 | | 2.4185 | H11 | 2.1662 | 2.3491 | 1.8894 | 0.9650 | 2.7221 | 2.9340 | 3.3839 | 2.6534 | 2.7707 | 2.4185 | |
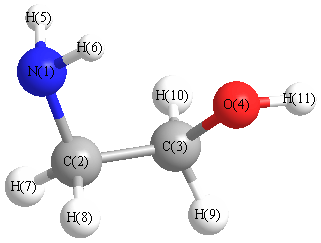 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.199 |
|
N1 |
C2 |
H7 |
114.275 |
| N1 |
C2 |
H8 |
107.735 |
|
C2 |
N1 |
H5 |
110.938 |
| C2 |
N1 |
H6 |
111.582 |
|
C2 |
C3 |
O4 |
110.454 |
| C2 |
C3 |
H9 |
110.393 |
|
C2 |
C3 |
H10 |
109.377 |
| C3 |
C2 |
H7 |
110.172 |
|
C3 |
C2 |
H8 |
108.517 |
| C3 |
O4 |
H11 |
104.375 |
|
O4 |
C3 |
H9 |
108.156 |
| O4 |
C3 |
H10 |
111.031 |
|
H5 |
N1 |
H6 |
107.264 |
| H7 |
C2 |
H8 |
107.785 |
|
H9 |
C3 |
H10 |
107.378 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.369 |
|
|
|
| 2 |
C |
-0.163 |
|
|
|
| 3 |
C |
0.007 |
|
|
|
| 4 |
O |
-0.365 |
|
|
|
| 5 |
H |
0.155 |
|
|
|
| 6 |
H |
0.157 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.094 |
|
|
|
| 10 |
H |
0.062 |
|
|
|
| 11 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.826 |
1.032 |
0.524 |
3.054 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.177 |
2.109 |
0.318 |
| y |
2.109 |
-25.419 |
-0.262 |
| z |
0.318 |
-0.262 |
-24.442 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.247 |
2.109 |
0.318 |
| y |
2.109 |
-0.109 |
-0.262 |
| z |
0.318 |
-0.262 |
1.356 |
|
| Polar |
|---|
| 3z2-r2 | 2.711 |
|---|
| x2-y2 | -0.758 |
|---|
| xy | 2.109 |
|---|
| xz | 0.318 |
|---|
| yz | -0.262 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.030 |
0.191 |
0.036 |
| y |
0.191 |
5.698 |
0.050 |
| z |
0.036 |
0.050 |
5.213 |
<r2> (average value of r
2) Å
2
| <r2> |
85.695 |
| (<r2>)1/2 |
9.257 |
