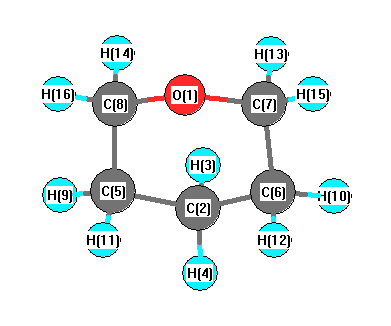
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3111 |
2977 |
38.09 |
|
|
|
| 2 |
A' |
3103 |
2970 |
52.08 |
|
|
|
| 3 |
A' |
3093 |
2960 |
39.70 |
|
|
|
| 4 |
A' |
3052 |
2921 |
18.75 |
|
|
|
| 5 |
A' |
3036 |
2906 |
23.50 |
|
|
|
| 6 |
A' |
2969 |
2842 |
104.78 |
|
|
|
| 7 |
A' |
1501 |
1436 |
3.82 |
|
|
|
| 8 |
A' |
1488 |
1424 |
7.20 |
|
|
|
| 9 |
A' |
1479 |
1415 |
7.94 |
|
|
|
| 10 |
A' |
1420 |
1359 |
6.09 |
|
|
|
| 11 |
A' |
1381 |
1322 |
2.34 |
|
|
|
| 12 |
A' |
1333 |
1276 |
5.83 |
|
|
|
| 13 |
A' |
1285 |
1230 |
5.01 |
|
|
|
| 14 |
A' |
1178 |
1127 |
3.38 |
|
|
|
| 15 |
A' |
1064 |
1019 |
10.73 |
|
|
|
| 16 |
A' |
1030 |
986 |
9.34 |
|
|
|
| 17 |
A' |
909 |
870 |
21.06 |
|
|
|
| 18 |
A' |
860 |
823 |
4.92 |
|
|
|
| 19 |
A' |
844 |
808 |
1.62 |
|
|
|
| 20 |
A' |
545 |
522 |
4.13 |
|
|
|
| 21 |
A' |
427 |
409 |
1.36 |
|
|
|
| 22 |
A' |
389 |
372 |
3.64 |
|
|
|
| 23 |
A' |
247 |
236 |
3.32 |
|
|
|
| 24 |
A" |
3108 |
2975 |
46.55 |
|
|
|
| 25 |
A" |
3098 |
2965 |
42.63 |
|
|
|
| 26 |
A" |
3053 |
2922 |
26.76 |
|
|
|
| 27 |
A" |
2964 |
2837 |
20.47 |
|
|
|
| 28 |
A" |
1489 |
1425 |
1.38 |
|
|
|
| 29 |
A" |
1469 |
1406 |
2.47 |
|
|
|
| 30 |
A" |
1395 |
1335 |
4.19 |
|
|
|
| 31 |
A" |
1379 |
1320 |
1.18 |
|
|
|
| 32 |
A" |
1354 |
1296 |
0.00 |
|
|
|
| 33 |
A" |
1306 |
1250 |
8.24 |
|
|
|
| 34 |
A" |
1240 |
1186 |
54.15 |
|
|
|
| 35 |
A" |
1193 |
1142 |
0.03 |
|
|
|
| 36 |
A" |
1148 |
1098 |
63.95 |
|
|
|
| 37 |
A" |
1073 |
1027 |
11.14 |
|
|
|
| 38 |
A" |
980 |
938 |
2.73 |
|
|
|
| 39 |
A" |
904 |
865 |
0.31 |
|
|
|
| 40 |
A" |
817 |
782 |
0.43 |
|
|
|
| 41 |
A" |
456 |
437 |
0.02 |
|
|
|
| 42 |
A" |
235 |
225 |
0.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32203.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30821.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.278 |
|
|
|
| 2 |
C |
-0.243 |
|
|
|
| 3 |
H |
0.124 |
|
|
|
| 4 |
H |
0.119 |
|
|
|
| 5 |
C |
-0.222 |
|
|
|
| 6 |
C |
-0.222 |
|
|
|
| 7 |
C |
-0.059 |
|
|
|
| 8 |
C |
-0.059 |
|
|
|
| 9 |
H |
0.112 |
|
|
|
| 10 |
H |
0.112 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.084 |
|
|
|
| 14 |
H |
0.084 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
H |
0.110 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.935 |
1.030 |
0.000 |
1.391 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.967 |
-1.563 |
0.000 |
| y |
-1.563 |
-40.115 |
0.000 |
| z |
0.000 |
0.000 |
-35.511 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.154 |
-1.563 |
0.000 |
| y |
-1.563 |
-2.876 |
0.000 |
| z |
0.000 |
0.000 |
4.030 |
|
| Polar |
|---|
| 3z2-r2 | 8.060 |
|---|
| x2-y2 | 1.148 |
|---|
| xy | -1.563 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.223 |
0.096 |
0.000 |
| y |
0.096 |
8.972 |
0.000 |
| z |
0.000 |
0.000 |
9.818 |
<r2> (average value of r
2) Å
2
| <r2> |
150.295 |
| (<r2>)1/2 |
12.259 |