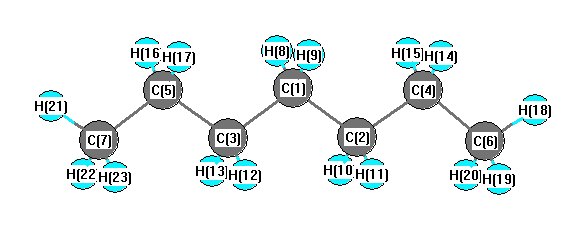
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3125 |
2991 |
46.17 |
|
|
|
| 2 |
A1 |
3050 |
2919 |
49.72 |
|
|
|
| 3 |
A1 |
3045 |
2914 |
119.07 |
|
|
|
| 4 |
A1 |
3034 |
2903 |
11.25 |
|
|
|
| 5 |
A1 |
3025 |
2896 |
0.37 |
|
|
|
| 6 |
A1 |
1514 |
1449 |
11.89 |
|
|
|
| 7 |
A1 |
1502 |
1438 |
0.48 |
|
|
|
| 8 |
A1 |
1489 |
1425 |
0.05 |
|
|
|
| 9 |
A1 |
1484 |
1420 |
0.00 |
|
|
|
| 10 |
A1 |
1410 |
1349 |
3.62 |
|
|
|
| 11 |
A1 |
1399 |
1339 |
0.06 |
|
|
|
| 12 |
A1 |
1311 |
1255 |
0.04 |
|
|
|
| 13 |
A1 |
1167 |
1117 |
0.70 |
|
|
|
| 14 |
A1 |
1088 |
1041 |
0.78 |
|
|
|
| 15 |
A1 |
1019 |
975 |
0.00 |
|
|
|
| 16 |
A1 |
921 |
882 |
1.36 |
|
|
|
| 17 |
A1 |
403 |
386 |
0.03 |
|
|
|
| 18 |
A1 |
302 |
289 |
0.00 |
|
|
|
| 19 |
A1 |
93 |
89 |
0.00 |
|
|
|
| 20 |
A2 |
3119 |
2985 |
0.00 |
|
|
|
| 21 |
A2 |
3080 |
2948 |
0.00 |
|
|
|
| 22 |
A2 |
3054 |
2923 |
0.00 |
|
|
|
| 23 |
A2 |
1497 |
1433 |
0.00 |
|
|
|
| 24 |
A2 |
1329 |
1272 |
0.00 |
|
|
|
| 25 |
A2 |
1325 |
1268 |
0.00 |
|
|
|
| 26 |
A2 |
1241 |
1187 |
0.00 |
|
|
|
| 27 |
A2 |
1025 |
981 |
0.00 |
|
|
|
| 28 |
A2 |
841 |
805 |
0.00 |
|
|
|
| 29 |
A2 |
734 |
702 |
0.00 |
|
|
|
| 30 |
A2 |
238 |
228 |
0.00 |
|
|
|
| 31 |
A2 |
141 |
135 |
0.00 |
|
|
|
| 32 |
A2 |
83 |
79 |
0.00 |
|
|
|
| 33 |
B1 |
3119 |
2985 |
120.70 |
|
|
|
| 34 |
B1 |
3090 |
2957 |
98.49 |
|
|
|
| 35 |
B1 |
3067 |
2936 |
0.01 |
|
|
|
| 36 |
B1 |
3046 |
2915 |
0.08 |
|
|
|
| 37 |
B1 |
1497 |
1433 |
14.59 |
|
|
|
| 38 |
B1 |
1336 |
1278 |
0.87 |
|
|
|
| 39 |
B1 |
1291 |
1235 |
0.16 |
|
|
|
| 40 |
B1 |
1205 |
1153 |
0.04 |
|
|
|
| 41 |
B1 |
935 |
895 |
0.69 |
|
|
|
| 42 |
B1 |
767 |
735 |
1.25 |
|
|
|
| 43 |
B1 |
729 |
698 |
5.81 |
|
|
|
| 44 |
B1 |
239 |
229 |
0.00 |
|
|
|
| 45 |
B1 |
149 |
143 |
0.00 |
|
|
|
| 46 |
B1 |
65 |
62 |
0.01 |
|
|
|
| 47 |
B2 |
3125 |
2990 |
32.16 |
|
|
|
| 48 |
B2 |
3049 |
2919 |
47.86 |
|
|
|
| 49 |
B2 |
3042 |
2911 |
0.14 |
|
|
|
| 50 |
B2 |
3027 |
2897 |
0.00 |
|
|
|
| 51 |
B2 |
1509 |
1444 |
1.06 |
|
|
|
| 52 |
B2 |
1494 |
1430 |
1.16 |
|
|
|
| 53 |
B2 |
1484 |
1420 |
0.20 |
|
|
|
| 54 |
B2 |
1411 |
1351 |
1.00 |
|
|
|
| 55 |
B2 |
1406 |
1346 |
1.78 |
|
|
|
| 56 |
B2 |
1365 |
1306 |
1.32 |
|
|
|
| 57 |
B2 |
1253 |
1199 |
1.57 |
|
|
|
| 58 |
B2 |
1098 |
1051 |
1.97 |
|
|
|
| 59 |
B2 |
1084 |
1038 |
1.50 |
|
|
|
| 60 |
B2 |
1060 |
1015 |
0.08 |
|
|
|
| 61 |
B2 |
893 |
854 |
2.53 |
|
|
|
| 62 |
B2 |
473 |
453 |
0.05 |
|
|
|
| 63 |
B2 |
232 |
222 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 47809.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 45758.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.213 |
|
|
|
| 2 |
C |
-0.207 |
|
|
|
| 3 |
C |
-0.207 |
|
|
|
| 4 |
C |
-0.206 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
C |
-0.366 |
|
|
|
| 7 |
C |
-0.366 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.108 |
|
|
|
| 11 |
H |
0.108 |
|
|
|
| 12 |
H |
0.108 |
|
|
|
| 13 |
H |
0.108 |
|
|
|
| 14 |
H |
0.110 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
H |
0.110 |
|
|
|
| 17 |
H |
0.110 |
|
|
|
| 18 |
H |
0.118 |
|
|
|
| 19 |
H |
0.111 |
|
|
|
| 20 |
H |
0.111 |
|
|
|
| 21 |
H |
0.118 |
|
|
|
| 22 |
H |
0.111 |
|
|
|
| 23 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.085 |
0.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.367 |
0.000 |
0.000 |
| y |
0.000 |
-49.658 |
0.000 |
| z |
0.000 |
0.000 |
-48.973 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.949 |
0.000 |
0.000 |
| y |
0.000 |
-1.488 |
0.000 |
| z |
0.000 |
0.000 |
-0.461 |
|
| Polar |
|---|
| 3z2-r2 | -0.921 |
|---|
| x2-y2 | 2.291 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.032 |
0.000 |
0.000 |
| y |
0.000 |
15.910 |
0.000 |
| z |
0.000 |
0.000 |
11.755 |
<r2> (average value of r
2) Å
2
| <r2> |
470.066 |
| (<r2>)1/2 |
21.681 |