Jump to
S1C2
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -255.141102 |
| Energy at 298.15K | -255.140488 |
| HF Energy | -255.141102 |
| Nuclear repulsion energy | 46.684614 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.691 |
| N2 |
0.000 |
0.000 |
-1.851 |
| Na3 |
0.000 |
0.000 |
1.555 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1596 | 2.2456 |
N2 | 1.1596 | | 3.4053 | Na3 | 2.2456 | 3.4053 | |
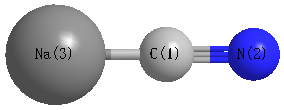 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.280 |
|
|
|
| 2 |
N |
-0.179 |
|
|
|
| 3 |
Na |
0.459 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.705 |
10.705 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.109 |
0.000 |
0.000 |
| y |
0.000 |
-17.109 |
0.000 |
| z |
0.000 |
0.000 |
-13.247 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.931 |
0.000 |
0.000 |
| y |
0.000 |
-1.931 |
0.000 |
| z |
0.000 |
0.000 |
3.863 |
|
| Polar |
|---|
| 3z2-r2 | 7.725 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.021 |
0.000 |
-0.000 |
| y |
0.000 |
3.022 |
-0.000 |
| z |
-0.000 |
-0.000 |
5.408 |
<r2> (average value of r
2) Å
2
| <r2> |
63.306 |
| (<r2>)1/2 |
7.957 |
Jump to
S1C1
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -255.145763 |
| Energy at 298.15K | -255.145525 |
| HF Energy | -255.145763 |
| Nuclear repulsion energy | 51.881214 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.078 |
0.646 |
0.000 |
| N2 |
0.000 |
1.103 |
0.000 |
| Na3 |
-0.588 |
-1.054 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1707 | 2.3805 |
N2 | 1.1707 | | 2.2357 | Na3 | 2.3805 | 2.2357 | |
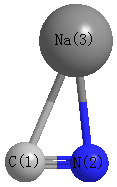 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
82.292 |
|
C1 |
Na3 |
N2 |
29.166 |
| N2 |
C1 |
Na3 |
68.542 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.172 |
|
|
|
| 2 |
N |
-0.204 |
|
|
|
| 3 |
Na |
0.376 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.582 |
-7.387 |
0.000 |
8.692 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.522 |
3.477 |
0.000 |
| y |
3.477 |
-13.910 |
0.000 |
| z |
0.000 |
0.000 |
-17.224 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.954 |
3.477 |
0.000 |
| y |
3.477 |
4.463 |
0.000 |
| z |
0.000 |
0.000 |
-0.508 |
|
| Polar |
|---|
| 3z2-r2 | -1.017 |
|---|
| x2-y2 | -5.612 |
|---|
| xy | 3.477 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.243 |
-0.070 |
0.000 |
| y |
-0.070 |
3.774 |
0.000 |
| z |
0.000 |
0.000 |
3.158 |
<r2> (average value of r
2) Å
2
| <r2> |
44.566 |
| (<r2>)1/2 |
6.676 |
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -255.141685 |
| Energy at 298.15K | -255.140803 |
| HF Energy | -255.141685 |
| Nuclear repulsion energy | 48.889341 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.847 |
| N2 |
0.000 |
0.000 |
-0.677 |
| Na3 |
0.000 |
0.000 |
1.438 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1701 | 3.2853 |
N2 | 1.1701 | | 2.1152 | Na3 | 3.2853 | 2.1152 | |
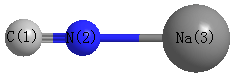 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.354 |
|
|
|
| 2 |
N |
-0.294 |
|
|
|
| 3 |
Na |
0.648 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.865 |
10.865 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.107 |
0.000 |
0.000 |
| y |
0.000 |
-17.107 |
0.000 |
| z |
0.000 |
0.000 |
-15.831 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.638 |
0.000 |
0.000 |
| y |
0.000 |
-0.638 |
0.000 |
| z |
0.000 |
0.000 |
1.277 |
|
| Polar |
|---|
| 3z2-r2 | 2.553 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.009 |
0.000 |
0.000 |
| y |
0.000 |
3.009 |
-0.000 |
| z |
0.000 |
-0.000 |
5.371 |
<r2> (average value of r
2) Å
2
| <r2> |
56.851 |
| (<r2>)1/2 |
7.540 |
