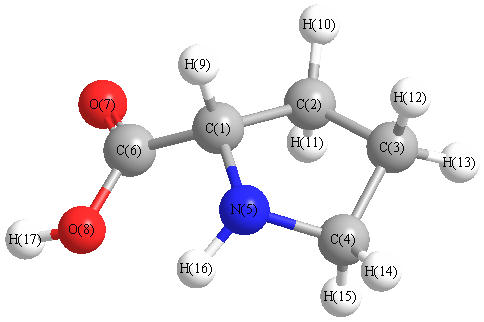
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3796 |
3633 |
73.14 |
|
|
|
| 2 |
A |
3594 |
3440 |
6.32 |
|
|
|
| 3 |
A |
3159 |
3024 |
18.64 |
|
|
|
| 4 |
A |
3134 |
3000 |
11.82 |
|
|
|
| 5 |
A |
3113 |
2979 |
23.82 |
|
|
|
| 6 |
A |
3107 |
2974 |
48.46 |
|
|
|
| 7 |
A |
3090 |
2957 |
17.12 |
|
|
|
| 8 |
A |
3057 |
2925 |
18.86 |
|
|
|
| 9 |
A |
2961 |
2834 |
72.19 |
|
|
|
| 10 |
A |
1850 |
1771 |
264.30 |
|
|
|
| 11 |
A |
1526 |
1461 |
0.50 |
|
|
|
| 12 |
A |
1507 |
1443 |
7.14 |
|
|
|
| 13 |
A |
1488 |
1424 |
0.89 |
|
|
|
| 14 |
A |
1457 |
1395 |
21.65 |
|
|
|
| 15 |
A |
1377 |
1318 |
22.85 |
|
|
|
| 16 |
A |
1371 |
1312 |
12.58 |
|
|
|
| 17 |
A |
1334 |
1277 |
7.06 |
|
|
|
| 18 |
A |
1328 |
1271 |
17.61 |
|
|
|
| 19 |
A |
1318 |
1261 |
3.34 |
|
|
|
| 20 |
A |
1256 |
1202 |
2.31 |
|
|
|
| 21 |
A |
1250 |
1196 |
2.65 |
|
|
|
| 22 |
A |
1222 |
1170 |
16.89 |
|
|
|
| 23 |
A |
1205 |
1153 |
7.95 |
|
|
|
| 24 |
A |
1171 |
1121 |
192.36 |
|
|
|
| 25 |
A |
1160 |
1110 |
51.32 |
|
|
|
| 26 |
A |
1108 |
1060 |
3.37 |
|
|
|
| 27 |
A |
1085 |
1039 |
4.27 |
|
|
|
| 28 |
A |
1012 |
969 |
4.59 |
|
|
|
| 29 |
A |
975 |
933 |
7.33 |
|
|
|
| 30 |
A |
944 |
903 |
5.37 |
|
|
|
| 31 |
A |
926 |
886 |
3.98 |
|
|
|
| 32 |
A |
902 |
863 |
65.06 |
|
|
|
| 33 |
A |
851 |
815 |
22.34 |
|
|
|
| 34 |
A |
784 |
750 |
2.00 |
|
|
|
| 35 |
A |
759 |
727 |
36.55 |
|
|
|
| 36 |
A |
678 |
649 |
92.59 |
|
|
|
| 37 |
A |
621 |
595 |
47.21 |
|
|
|
| 38 |
A |
595 |
569 |
10.91 |
|
|
|
| 39 |
A |
519 |
497 |
33.09 |
|
|
|
| 40 |
A |
497 |
476 |
13.22 |
|
|
|
| 41 |
A |
361 |
345 |
0.22 |
|
|
|
| 42 |
A |
248 |
238 |
3.64 |
|
|
|
| 43 |
A |
185 |
177 |
0.19 |
|
|
|
| 44 |
A |
67 |
64 |
0.07 |
|
|
|
| 45 |
A |
40 |
38 |
2.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31993.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30621.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.056 |
|
|
|
| 2 |
C |
-0.232 |
|
|
|
| 3 |
C |
-0.214 |
|
|
|
| 4 |
C |
-0.150 |
|
|
|
| 5 |
N |
-0.225 |
|
|
|
| 6 |
C |
0.265 |
|
|
|
| 7 |
O |
-0.306 |
|
|
|
| 8 |
O |
-0.270 |
|
|
|
| 9 |
H |
0.135 |
|
|
|
| 10 |
H |
0.123 |
|
|
|
| 11 |
H |
0.132 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.113 |
|
|
|
| 15 |
H |
0.094 |
|
|
|
| 16 |
H |
0.133 |
|
|
|
| 17 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.173 |
-1.404 |
-0.340 |
1.455 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.459 |
5.847 |
-1.108 |
| y |
5.847 |
-49.126 |
2.406 |
| z |
-1.108 |
2.406 |
-47.121 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.665 |
5.847 |
-1.108 |
| y |
5.847 |
-2.336 |
2.406 |
| z |
-1.108 |
2.406 |
0.671 |
|
| Polar |
|---|
| 3z2-r2 | 1.343 |
|---|
| x2-y2 | 2.667 |
|---|
| xy | 5.847 |
|---|
| xz | -1.108 |
|---|
| yz | 2.406 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.939 |
0.031 |
0.046 |
| y |
0.031 |
10.350 |
0.191 |
| z |
0.046 |
0.191 |
8.759 |
<r2> (average value of r
2) Å
2
| <r2> |
253.431 |
| (<r2>)1/2 |
15.920 |