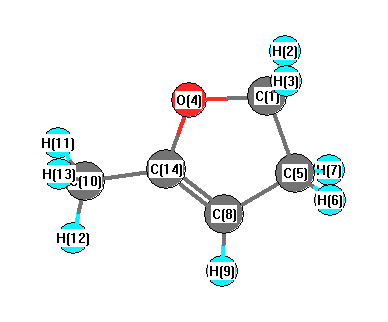
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3123 |
3.97 |
|
|
|
| 2 |
A |
3165 |
3029 |
11.96 |
|
|
|
| 3 |
A |
3133 |
2999 |
33.60 |
|
|
|
| 4 |
A |
3117 |
2984 |
11.01 |
|
|
|
| 5 |
A |
3094 |
2961 |
35.18 |
|
|
|
| 6 |
A |
3060 |
2929 |
20.72 |
|
|
|
| 7 |
A |
3051 |
2920 |
49.90 |
|
|
|
| 8 |
A |
3030 |
2900 |
48.67 |
|
|
|
| 9 |
A |
1753 |
1678 |
62.67 |
|
|
|
| 10 |
A |
1521 |
1455 |
1.43 |
|
|
|
| 11 |
A |
1497 |
1433 |
1.10 |
|
|
|
| 12 |
A |
1487 |
1423 |
4.85 |
|
|
|
| 13 |
A |
1470 |
1407 |
7.61 |
|
|
|
| 14 |
A |
1424 |
1363 |
31.92 |
|
|
|
| 15 |
A |
1403 |
1343 |
2.01 |
|
|
|
| 16 |
A |
1331 |
1274 |
7.41 |
|
|
|
| 17 |
A |
1277 |
1223 |
55.06 |
|
|
|
| 18 |
A |
1265 |
1211 |
7.84 |
|
|
|
| 19 |
A |
1218 |
1166 |
33.30 |
|
|
|
| 20 |
A |
1213 |
1161 |
5.11 |
|
|
|
| 21 |
A |
1082 |
1035 |
0.33 |
|
|
|
| 22 |
A |
1064 |
1018 |
28.48 |
|
|
|
| 23 |
A |
1059 |
1014 |
28.71 |
|
|
|
| 24 |
A |
1033 |
988 |
5.19 |
|
|
|
| 25 |
A |
993 |
950 |
12.29 |
|
|
|
| 26 |
A |
953 |
912 |
11.05 |
|
|
|
| 27 |
A |
929 |
890 |
28.48 |
|
|
|
| 28 |
A |
864 |
827 |
4.27 |
|
|
|
| 29 |
A |
743 |
712 |
28.99 |
|
|
|
| 30 |
A |
690 |
660 |
0.78 |
|
|
|
| 31 |
A |
638 |
611 |
0.95 |
|
|
|
| 32 |
A |
568 |
544 |
0.56 |
|
|
|
| 33 |
A |
334 |
320 |
2.34 |
|
|
|
| 34 |
A |
230 |
220 |
6.37 |
|
|
|
| 35 |
A |
177 |
170 |
2.44 |
|
|
|
| 36 |
A |
133 |
127 |
1.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26630.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25487.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.025 |
|
|
|
| 2 |
H |
0.100 |
|
|
|
| 3 |
H |
0.090 |
|
|
|
| 4 |
O |
-0.244 |
|
|
|
| 5 |
C |
-0.193 |
|
|
|
| 6 |
H |
0.110 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
C |
-0.309 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
| 10 |
C |
-0.350 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.125 |
|
|
|
| 14 |
C |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.356 |
0.621 |
0.130 |
0.728 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.338 |
1.132 |
-0.026 |
| y |
1.132 |
-37.507 |
-0.205 |
| z |
-0.026 |
-0.205 |
-37.209 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.020 |
1.132 |
-0.026 |
| y |
1.132 |
-2.734 |
-0.205 |
| z |
-0.026 |
-0.205 |
-2.287 |
|
| Polar |
|---|
| 3z2-r2 | -4.573 |
|---|
| x2-y2 | 5.169 |
|---|
| xy | 1.132 |
|---|
| xz | -0.026 |
|---|
| yz | -0.205 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.685 |
-0.530 |
0.017 |
| y |
-0.530 |
9.038 |
-0.018 |
| z |
0.017 |
-0.018 |
6.719 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |