return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-194.931498
Energy at 298.15K-194.939034
HF Energy-194.027024
Nuclear repulsion energy156.683853
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3285 3119 9.73      
2 A' 3231 3067 8.92      
3 A' 3215 3052 20.53      
4 A' 3198 3036 3.78      
5 A' 3189 3028 6.03      
6 A' 3177 3017 0.80      
7 A' 3086 2930 19.86      
8 A' 1731 1643 5.29      
9 A' 1663 1579 2.36      
10 A' 1515 1439 7.47      
11 A' 1489 1413 12.44      
12 A' 1429 1357 3.60      
13 A' 1404 1333 1.21      
14 A' 1327 1260 0.64      
15 A' 1298 1233 1.39      
16 A' 1199 1138 0.95      
17 A' 1073 1019 0.04      
18 A' 982 932 10.03      
19 A' 912 866 2.22      
20 A' 616 584 4.47      
21 A' 388 368 3.38      
22 A' 224 213 0.21      
23 A" 3140 2981 11.52      
24 A" 1509 1432 6.34      
25 A" 1074 1019 2.55      
26 A" 1054 1001 16.48      
27 A" 1007 956 1.52      
28 A" 950 902 40.43      
29 A" 828 786 7.82      
30 A" 659 625 26.58      
31 A" 366 347 0.26      
32 A" 149 141 0.03      
33 A" 124 118 0.70      

Unscaled Zero Point Vibrational Energy (zpe) 25243.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23966.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.53412 0.08973 0.07791

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.374 0.149 0.000
C2 1.087 -0.212 0.000
C3 0.000 0.740 0.000
C4 -1.308 0.451 0.000
C5 -1.926 -0.903 0.000
H6 2.655 1.190 0.000
H7 3.163 -0.582 0.000
H8 0.846 -1.262 0.000
H9 0.288 1.780 0.000
H10 -1.997 1.282 0.000
H11 -1.191 -1.697 0.000
H12 -2.562 -1.031 0.873
H13 -2.562 -1.031 -0.873

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.33662.44633.69454.42701.07821.07582.07962.64774.51584.01485.14925.1492
C21.33661.44512.48563.09162.10362.10871.07692.14663.42752.71953.84003.8400
C32.44631.44511.33962.53142.69313.42832.17321.07952.06972.71253.23413.2341
C43.69452.48561.33961.48844.03164.58932.75272.07691.07922.15202.12872.1287
C54.42703.09162.53141.48845.03685.09972.79583.47862.18551.08281.08751.0875
H61.07822.10362.69314.03165.03681.84333.04702.43944.65354.80945.73675.7367
H71.07582.10873.42834.58935.09971.84332.41483.72095.48704.49495.80845.8084
H82.07961.07692.17322.75272.79583.04702.41483.09283.81542.08333.52553.5255
H92.64772.14661.07952.07693.47862.43943.72093.09282.33933.77894.09704.0970
H104.51583.42752.06971.07922.18554.65355.48703.81542.33933.08642.53582.5358
H114.01482.71952.71252.15201.08284.80944.49492.08333.77893.08641.75641.7564
H125.14923.84003.23412.12871.08755.73675.80843.52554.09702.53581.75641.7467
H135.14923.84003.23412.12871.08755.73675.80843.52554.09702.53581.75641.7467

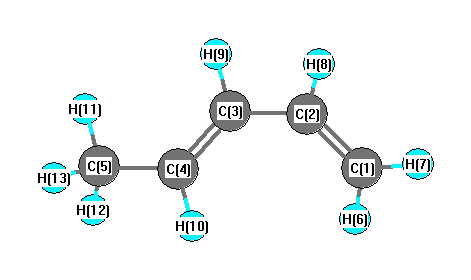
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.102 C1 C2 H8 118.616
C2 C1 H6 120.807 C2 C1 H7 121.509
C2 C3 C4 126.364 C2 C3 H9 115.726
C3 C2 H8 118.282 C3 C4 C5 126.968
C3 C4 H10 117.267 C4 C3 H9 117.911
C4 C5 H11 112.684 C4 C5 H12 110.491
C4 C5 H13 110.491 C5 C4 H10 115.765
H6 C1 H7 117.684 H11 C5 H12 108.053
H11 C5 H13 108.053 H12 C5 H13 106.852
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability