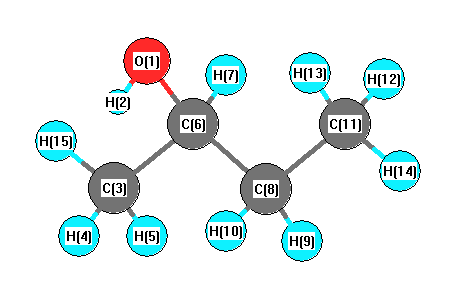
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3847 |
3682 |
16.93 |
|
|
|
| 2 |
A |
3144 |
3009 |
39.88 |
|
|
|
| 3 |
A |
3143 |
3008 |
5.99 |
|
|
|
| 4 |
A |
3126 |
2992 |
47.41 |
|
|
|
| 5 |
A |
3118 |
2984 |
31.71 |
|
|
|
| 6 |
A |
3078 |
2946 |
23.41 |
|
|
|
| 7 |
A |
3060 |
2929 |
36.48 |
|
|
|
| 8 |
A |
3058 |
2927 |
22.22 |
|
|
|
| 9 |
A |
3044 |
2913 |
14.18 |
|
|
|
| 10 |
A |
3024 |
2895 |
18.83 |
|
|
|
| 11 |
A |
1508 |
1443 |
6.78 |
|
|
|
| 12 |
A |
1497 |
1432 |
10.42 |
|
|
|
| 13 |
A |
1493 |
1429 |
0.69 |
|
|
|
| 14 |
A |
1491 |
1427 |
7.76 |
|
|
|
| 15 |
A |
1476 |
1413 |
0.32 |
|
|
|
| 16 |
A |
1419 |
1358 |
31.42 |
|
|
|
| 17 |
A |
1407 |
1347 |
17.37 |
|
|
|
| 18 |
A |
1399 |
1339 |
10.15 |
|
|
|
| 19 |
A |
1390 |
1331 |
2.07 |
|
|
|
| 20 |
A |
1335 |
1278 |
2.46 |
|
|
|
| 21 |
A |
1320 |
1263 |
3.79 |
|
|
|
| 22 |
A |
1258 |
1204 |
6.04 |
|
|
|
| 23 |
A |
1192 |
1141 |
22.57 |
|
|
|
| 24 |
A |
1144 |
1095 |
18.41 |
|
|
|
| 25 |
A |
1094 |
1047 |
74.06 |
|
|
|
| 26 |
A |
1060 |
1015 |
0.82 |
|
|
|
| 27 |
A |
1007 |
963 |
18.98 |
|
|
|
| 28 |
A |
976 |
934 |
8.84 |
|
|
|
| 29 |
A |
931 |
892 |
8.91 |
|
|
|
| 30 |
A |
821 |
786 |
5.08 |
|
|
|
| 31 |
A |
776 |
743 |
0.51 |
|
|
|
| 32 |
A |
492 |
471 |
12.13 |
|
|
|
| 33 |
A |
453 |
433 |
5.57 |
|
|
|
| 34 |
A |
369 |
353 |
8.93 |
|
|
|
| 35 |
A |
262 |
251 |
90.92 |
|
|
|
| 36 |
A |
246 |
235 |
6.31 |
|
|
|
| 37 |
A |
231 |
221 |
3.01 |
|
|
|
| 38 |
A |
218 |
209 |
6.07 |
|
|
|
| 39 |
A |
119 |
114 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30012.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28724.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.352 |
|
|
|
| 2 |
H |
0.206 |
|
|
|
| 3 |
C |
-0.356 |
|
|
|
| 4 |
H |
0.102 |
|
|
|
| 5 |
H |
0.108 |
|
|
|
| 6 |
C |
0.070 |
|
|
|
| 7 |
H |
0.107 |
|
|
|
| 8 |
C |
-0.215 |
|
|
|
| 9 |
H |
0.108 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
| 11 |
C |
-0.359 |
|
|
|
| 12 |
H |
0.111 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.115 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.186 |
-1.233 |
0.886 |
1.529 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.633 |
0.172 |
-0.640 |
| y |
0.172 |
-35.535 |
2.290 |
| z |
-0.640 |
2.290 |
-30.902 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.415 |
0.172 |
-0.640 |
| y |
0.172 |
-3.268 |
2.290 |
| z |
-0.640 |
2.290 |
3.682 |
|
| Polar |
|---|
| 3z2-r2 | 7.365 |
|---|
| x2-y2 | 1.902 |
|---|
| xy | 0.172 |
|---|
| xz | -0.640 |
|---|
| yz | 2.290 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.942 |
0.172 |
-0.077 |
| y |
0.172 |
7.648 |
0.063 |
| z |
-0.077 |
0.063 |
7.445 |
<r2> (average value of r
2) Å
2
| <r2> |
144.575 |
| (<r2>)1/2 |
12.024 |