return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-593.939107
Energy at 298.15K-593.951538
HF Energy-592.782123
Nuclear repulsion energy314.595018
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3142 2983 11.82      
2 A' 3126 2968 37.97      
3 A' 3118 2960 37.34      
4 A' 3077 2921 9.91      
5 A' 3075 2920 27.64      
6 A' 3063 2908 18.25      
7 A' 1513 1437 2.46      
8 A' 1500 1424 13.24      
9 A' 1483 1408 7.51      
10 A' 1393 1322 0.35      
11 A' 1352 1284 2.59      
12 A' 1284 1219 0.62      
13 A' 1259 1195 4.67      
14 A' 1097 1041 1.33      
15 A' 1056 1002 0.57      
16 A' 991 941 7.51      
17 A' 851 808 4.28      
18 A' 840 798 0.38      
19 A' 691 656 2.56      
20 A' 499 473 0.38      
21 A' 374 355 0.32      
22 A' 344 327 0.16      
23 A' 185 176 1.62      
24 A" 3144 2985 9.63      
25 A" 3120 2962 18.57      
26 A" 3080 2924 0.16      
27 A" 3078 2922 47.64      
28 A" 1494 1418 3.96      
29 A" 1479 1404 1.21      
30 A" 1400 1329 0.00      
31 A" 1384 1314 0.12      
32 A" 1314 1247 8.81      
33 A" 1307 1241 7.96      
34 A" 1182 1122 0.16      
35 A" 1130 1073 0.09      
36 A" 1112 1055 0.52      
37 A" 970 921 0.77      
38 A" 923 877 3.00      
39 A" 805 765 0.01      
40 A" 719 683 0.07      
41 A" 397 377 0.23      
42 A" 246 234 0.44      

Unscaled Zero Point Vibrational Energy (zpe) 31797.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 30188.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.13557 0.10158 0.06495

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -0.776 -1.279 0.000
C2 0.821 1.497 0.000
C3 0.149 0.995 1.266
C4 0.149 0.995 -1.266
C5 0.149 -0.517 1.351
C6 0.149 -0.517 -1.351
H7 0.827 2.584 0.000
H8 1.862 1.174 0.000
H9 1.167 -0.899 1.350
H10 1.167 -0.899 -1.350
H11 0.669 1.395 2.134
H12 0.669 1.395 -2.134
H13 -0.879 1.351 1.302
H14 -0.879 1.351 -1.302
H15 -0.337 -0.857 -2.260
H16 -0.337 -0.857 2.260

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.20232.76182.76181.80571.80574.18173.60242.39672.39673.71393.71392.93652.93652.34022.3402
C23.20231.51871.51872.51672.51671.08661.09012.77222.77222.14202.14202.14602.14603.46223.4622
C32.76181.51872.53181.51453.02242.14152.13802.15253.38691.08843.46281.08782.78864.01172.1569
C42.76181.51872.53183.02241.51452.14152.13803.38692.15253.46281.08842.78861.08782.15694.0117
C51.80572.51671.51453.02242.70223.44952.76081.08812.91232.13104.00942.13273.40383.65911.0849
C61.80572.51673.02241.51452.70223.44952.76082.91231.08814.00942.13103.40382.13271.08493.6591
H74.18171.08662.14152.14153.44953.44951.74883.75083.75082.44772.44772.47442.47444.27744.2774
H83.60241.09012.13802.13802.76082.76081.74882.57002.57002.45532.45533.03953.03953.75073.7507
H92.39672.77222.15253.38691.08812.91233.75082.57002.70072.47574.20193.04194.03563.91131.7586
H102.39672.77223.38692.15252.91231.08813.75082.57002.70074.20192.47574.03563.04191.75863.9113
H113.71392.14201.08843.46282.13104.00942.44772.45532.47574.20194.26831.75733.76885.03882.4697
H123.71392.14203.46281.08844.00942.13102.44772.45534.20192.47574.26833.76881.75732.46975.0388
H132.93652.14601.08782.78862.13273.40382.47443.03953.04194.03561.75733.76882.60424.22542.4667
H142.93652.14602.78861.08783.40382.13272.47443.03954.03563.04193.76881.75732.60422.46674.2254
H152.34023.46224.01172.15693.65911.08494.27743.75073.91131.75865.03882.46974.22542.46674.5193
H162.34023.46222.15694.01171.08493.65914.27743.75071.75863.91132.46975.03882.46674.22544.5193

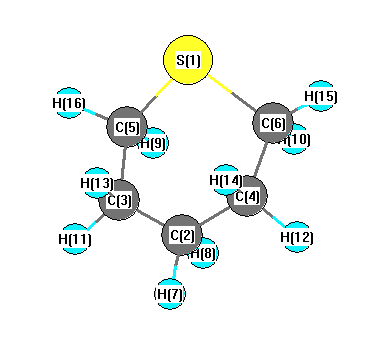
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.273 S1 C5 H9 109.313
S1 C5 H16 105.380 S1 C6 C4 112.273
S1 C6 H10 109.313 S1 C6 H15 105.380
C2 C3 C5 112.137 C2 C3 H11 109.377
C2 C3 H13 109.731 C2 C4 C6 112.137
C2 C4 H12 109.377 C2 C4 H14 109.731
C3 C2 C4 112.922 C3 C2 H7 109.443
C3 C2 H8 108.969 C3 C5 H9 110.531
C3 C5 H16 111.077 C4 C2 H7 109.443
C4 C2 H8 108.969 C4 C6 H10 110.531
C4 C6 H15 111.077 C5 S1 C6 96.875
C5 C3 H11 108.808 C5 C3 H13 108.978
C6 C4 H12 108.808 C6 C4 H14 108.978
H7 C2 H8 106.913 H9 C5 H16 108.063
H10 C6 H15 108.063 H11 C3 H13 107.702
H12 C4 H14 107.702
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability