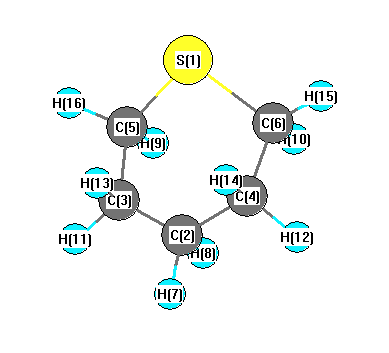
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3122 |
2988 |
18.20 |
|
|
|
| 2 |
A |
3096 |
2964 |
56.45 |
|
|
|
| 3 |
A |
3087 |
2954 |
46.11 |
|
|
|
| 4 |
A |
3049 |
2919 |
20.53 |
|
|
|
| 5 |
A |
3043 |
2912 |
24.72 |
|
|
|
| 6 |
A |
3030 |
2900 |
20.98 |
|
|
|
| 7 |
A |
1489 |
1425 |
3.57 |
|
|
|
| 8 |
A |
1475 |
1412 |
11.67 |
|
|
|
| 9 |
A |
1462 |
1400 |
8.70 |
|
|
|
| 10 |
A |
1383 |
1324 |
0.55 |
|
|
|
| 11 |
A |
1335 |
1278 |
2.70 |
|
|
|
| 12 |
A |
1267 |
1212 |
0.51 |
|
|
|
| 13 |
A |
1241 |
1187 |
5.01 |
|
|
|
| 14 |
A |
1080 |
1034 |
1.48 |
|
|
|
| 15 |
A |
1039 |
994 |
0.54 |
|
|
|
| 16 |
A |
981 |
939 |
7.43 |
|
|
|
| 17 |
A |
840 |
804 |
4.53 |
|
|
|
| 18 |
A |
829 |
793 |
0.52 |
|
|
|
| 19 |
A |
678 |
649 |
3.13 |
|
|
|
| 20 |
A |
478 |
457 |
0.46 |
|
|
|
| 21 |
A |
348 |
333 |
0.69 |
|
|
|
| 22 |
A |
332 |
318 |
0.00 |
|
|
|
| 23 |
A |
174 |
166 |
1.73 |
|
|
|
| 24 |
A |
3123 |
2989 |
11.43 |
|
|
|
| 25 |
A |
3089 |
2956 |
20.74 |
|
|
|
| 26 |
A |
3052 |
2921 |
15.57 |
|
|
|
| 27 |
A |
3044 |
2913 |
48.03 |
|
|
|
| 28 |
A |
1469 |
1406 |
4.26 |
|
|
|
| 29 |
A |
1459 |
1396 |
1.40 |
|
|
|
| 30 |
A |
1390 |
1330 |
0.01 |
|
|
|
| 31 |
A |
1371 |
1312 |
0.11 |
|
|
|
| 32 |
A |
1298 |
1242 |
9.48 |
|
|
|
| 33 |
A |
1289 |
1234 |
8.03 |
|
|
|
| 34 |
A |
1165 |
1115 |
0.10 |
|
|
|
| 35 |
A |
1110 |
1063 |
0.10 |
|
|
|
| 36 |
A |
1093 |
1046 |
0.58 |
|
|
|
| 37 |
A |
955 |
914 |
0.91 |
|
|
|
| 38 |
A |
909 |
870 |
3.32 |
|
|
|
| 39 |
A |
795 |
761 |
0.05 |
|
|
|
| 40 |
A |
708 |
677 |
0.16 |
|
|
|
| 41 |
A |
394 |
377 |
0.18 |
|
|
|
| 42 |
A |
230 |
220 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31398.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30051.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.105 |
|
|
|
| 2 |
C |
-0.231 |
|
|
|
| 3 |
C |
-0.228 |
|
|
|
| 4 |
C |
-0.228 |
|
|
|
| 5 |
C |
-0.238 |
|
|
|
| 6 |
C |
-0.238 |
|
|
|
| 7 |
H |
0.122 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.133 |
|
|
|
| 11 |
H |
0.119 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.128 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.133 |
|
|
|
| 16 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.222 |
1.335 |
0.000 |
1.810 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.831 |
-1.248 |
0.000 |
| y |
-1.248 |
-48.269 |
0.000 |
| z |
0.000 |
0.000 |
-42.058 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.667 |
-1.248 |
0.000 |
| y |
-1.248 |
-3.824 |
0.000 |
| z |
0.000 |
0.000 |
5.492 |
|
| Polar |
|---|
| 3z2-r2 | 10.983 |
|---|
| x2-y2 | 1.438 |
|---|
| xy | -1.248 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.865 |
1.051 |
0.000 |
| y |
1.051 |
11.202 |
0.000 |
| z |
0.000 |
0.000 |
12.017 |
<r2> (average value of r
2) Å
2
| <r2> |
194.113 |
| (<r2>)1/2 |
13.932 |