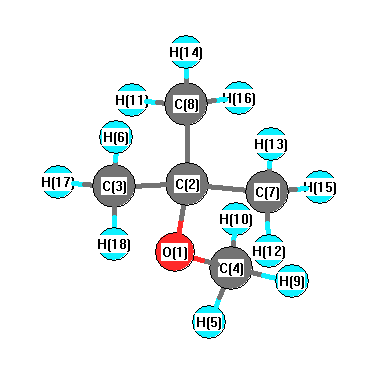
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3011 |
47.50 |
|
|
|
| 2 |
A |
3143 |
3009 |
30.27 |
|
|
|
| 3 |
A |
3138 |
3004 |
11.10 |
|
|
|
| 4 |
A |
3137 |
3003 |
9.49 |
|
|
|
| 5 |
A |
3136 |
3002 |
33.40 |
|
|
|
| 6 |
A |
3131 |
2997 |
2.75 |
|
|
|
| 7 |
A |
3130 |
2995 |
39.10 |
|
|
|
| 8 |
A |
3067 |
2935 |
74.24 |
|
|
|
| 9 |
A |
3065 |
2933 |
5.67 |
|
|
|
| 10 |
A |
3059 |
2928 |
26.12 |
|
|
|
| 11 |
A |
3054 |
2923 |
1.04 |
|
|
|
| 12 |
A |
3012 |
2883 |
47.08 |
|
|
|
| 13 |
A |
1526 |
1460 |
5.00 |
|
|
|
| 14 |
A |
1518 |
1453 |
10.37 |
|
|
|
| 15 |
A |
1513 |
1449 |
4.13 |
|
|
|
| 16 |
A |
1500 |
1435 |
2.29 |
|
|
|
| 17 |
A |
1492 |
1428 |
4.31 |
|
|
|
| 18 |
A |
1486 |
1423 |
1.40 |
|
|
|
| 19 |
A |
1483 |
1419 |
0.02 |
|
|
|
| 20 |
A |
1476 |
1413 |
0.55 |
|
|
|
| 21 |
A |
1466 |
1404 |
0.01 |
|
|
|
| 22 |
A |
1414 |
1353 |
14.50 |
|
|
|
| 23 |
A |
1394 |
1334 |
20.64 |
|
|
|
| 24 |
A |
1387 |
1328 |
22.86 |
|
|
|
| 25 |
A |
1295 |
1240 |
12.48 |
|
|
|
| 26 |
A |
1262 |
1208 |
17.36 |
|
|
|
| 27 |
A |
1243 |
1190 |
101.61 |
|
|
|
| 28 |
A |
1210 |
1158 |
6.53 |
|
|
|
| 29 |
A |
1186 |
1135 |
1.50 |
|
|
|
| 30 |
A |
1150 |
1101 |
92.79 |
|
|
|
| 31 |
A |
1041 |
996 |
1.51 |
|
|
|
| 32 |
A |
1033 |
989 |
2.14 |
|
|
|
| 33 |
A |
949 |
909 |
0.09 |
|
|
|
| 34 |
A |
925 |
885 |
0.15 |
|
|
|
| 35 |
A |
915 |
876 |
0.01 |
|
|
|
| 36 |
A |
879 |
841 |
18.20 |
|
|
|
| 37 |
A |
743 |
711 |
4.61 |
|
|
|
| 38 |
A |
503 |
482 |
1.73 |
|
|
|
| 39 |
A |
450 |
431 |
6.41 |
|
|
|
| 40 |
A |
400 |
383 |
0.26 |
|
|
|
| 41 |
A |
367 |
351 |
1.03 |
|
|
|
| 42 |
A |
334 |
320 |
0.21 |
|
|
|
| 43 |
A |
309 |
296 |
2.44 |
|
|
|
| 44 |
A |
287 |
275 |
0.91 |
|
|
|
| 45 |
A |
250 |
239 |
0.22 |
|
|
|
| 46 |
A |
238 |
227 |
0.61 |
|
|
|
| 47 |
A |
183 |
175 |
0.58 |
|
|
|
| 48 |
A |
35 |
34 |
2.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36031.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34485.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.275 |
|
|
|
| 2 |
C |
0.176 |
|
|
|
| 3 |
C |
-0.338 |
|
|
|
| 4 |
C |
-0.183 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.104 |
|
|
|
| 7 |
C |
-0.383 |
|
|
|
| 8 |
C |
-0.384 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
| 11 |
H |
0.131 |
|
|
|
| 12 |
H |
0.131 |
|
|
|
| 13 |
H |
0.120 |
|
|
|
| 14 |
H |
0.120 |
|
|
|
| 15 |
H |
0.124 |
|
|
|
| 16 |
H |
0.124 |
|
|
|
| 17 |
H |
0.119 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.050 |
-0.002 |
1.170 |
1.171 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.332 |
-0.007 |
-0.510 |
| y |
-0.007 |
-39.870 |
-0.009 |
| z |
-0.510 |
-0.009 |
-41.292 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.249 |
-0.007 |
-0.510 |
| y |
-0.007 |
-0.558 |
-0.009 |
| z |
-0.510 |
-0.009 |
-2.691 |
|
| Polar |
|---|
| 3z2-r2 | -5.381 |
|---|
| x2-y2 | 2.538 |
|---|
| xy | -0.007 |
|---|
| xz | -0.510 |
|---|
| yz | -0.009 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.562 |
0.002 |
-0.100 |
| y |
0.002 |
9.440 |
-0.001 |
| z |
-0.100 |
-0.001 |
9.225 |
<r2> (average value of r
2) Å
2
| <r2> |
177.559 |
| (<r2>)1/2 |
13.325 |