return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH2 (methylallyl radical)

using model chemistry: MP4=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 2A"
Energy calculated at MP4=FULL/cc-pVTZ
 hartrees
Energy at 0K-156.334436
Energy at 298.15K-156.340894
HF Energy-155.562650
Nuclear repulsion energy109.846393
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3266 3145        
2 A' 3184 3066        
3 A' 3177 3059        
4 A' 3164 3047        
5 A' 3117 3001        
6 A' 3035 2922        
7 A' 1540 1483        
8 A' 1527 1470        
9 A' 1492 1437        
10 A' 1419 1367        
11 A' 1356 1306        
12 A' 1309 1261        
13 A' 1172 1129        
14 A' 1146 1103        
15 A' 997 960        
16 A' 893 860        
17 A' 507 488        
18 A' 281 271        
19 A" 3077 2963        
20 A" 1497 1441        
21 A" 1073 1033        
22 A" 1044 1005        
23 A" 845 814        
24 A" 787 757        
25 A" 575 554        
26 A" 241 233        
27 A" 166 160        

Unscaled Zero Point Vibrational Energy (zpe) 20944.3 cm-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 20167.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVTZ
ABC
1.28680 0.13529 0.12523

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.356 -1.400 0.000
C2 0.150 -0.738 0.000
C3 0.000 0.625 0.000
C4 -1.314 1.326 0.000
H5 2.286 -0.854 0.000
H6 1.405 -2.476 0.000
H7 -0.752 -1.334 0.000
H8 0.890 1.238 0.000
H9 -2.137 0.614 0.000
H10 -1.421 1.969 0.875
H11 -1.421 1.969 -0.875

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
C11.37552.43763.81591.07891.07652.10932.67944.03254.45304.4530
C21.37551.37182.53072.13912.14351.08172.11062.65743.25003.2500
C32.43761.37181.48922.72303.40472.09911.08092.13742.14262.1426
C43.81592.53071.48924.20884.67412.71862.20641.08801.09141.0914
H51.07892.13912.72304.20881.84553.07632.51514.66084.74114.7411
H61.07652.14353.40474.67411.84552.44123.74964.70105.33945.3394
H72.10931.08172.09912.71863.07632.44123.05232.39053.48173.4817
H82.67942.11061.08092.20642.51513.74963.05233.09142.57752.5775
H94.03252.65742.13741.08804.66084.70102.39053.09141.76451.7645
H104.45303.25002.14261.09144.74115.33943.48172.57751.76451.7510
H114.45303.25002.14261.09144.74115.33943.48172.57751.76451.7510

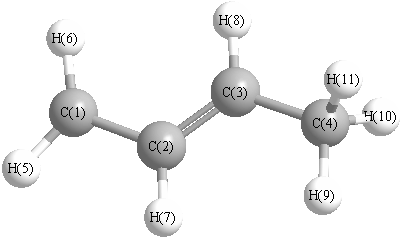
picture of methylallyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.064 C1 C2 H7 117.784
C2 C1 H5 120.793 C2 C1 H6 121.421
C2 C3 C4 124.337 C2 C3 H8 118.260
C3 C2 H7 117.151 C3 C4 H9 111.107
C3 C4 H10 111.318 C3 C4 H11 111.318
C4 C3 H8 117.403 H5 C1 H6 117.787
H9 C4 H10 108.119 H9 C4 H11 108.119
H10 C4 H11 106.672
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability