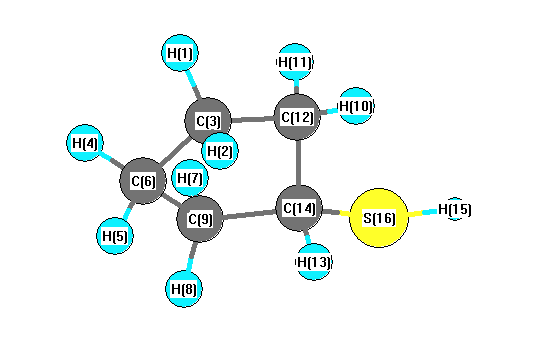
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3138 |
3003 |
61.52 |
|
|
|
| 2 |
A |
3125 |
2991 |
19.09 |
|
|
|
| 3 |
A |
3121 |
2987 |
8.79 |
|
|
|
| 4 |
A |
3108 |
2975 |
21.88 |
|
|
|
| 5 |
A |
3086 |
2954 |
38.74 |
|
|
|
| 6 |
A |
3077 |
2945 |
30.52 |
|
|
|
| 7 |
A |
3069 |
2938 |
18.02 |
|
|
|
| 8 |
A |
3064 |
2932 |
14.38 |
|
|
|
| 9 |
A |
3059 |
2928 |
8.32 |
|
|
|
| 10 |
A |
2712 |
2595 |
4.73 |
|
|
|
| 11 |
A |
1518 |
1453 |
1.32 |
|
|
|
| 12 |
A |
1495 |
1431 |
6.08 |
|
|
|
| 13 |
A |
1492 |
1428 |
3.41 |
|
|
|
| 14 |
A |
1484 |
1420 |
1.11 |
|
|
|
| 15 |
A |
1356 |
1298 |
3.97 |
|
|
|
| 16 |
A |
1347 |
1289 |
0.73 |
|
|
|
| 17 |
A |
1335 |
1278 |
1.06 |
|
|
|
| 18 |
A |
1324 |
1268 |
2.29 |
|
|
|
| 19 |
A |
1298 |
1243 |
0.26 |
|
|
|
| 20 |
A |
1272 |
1217 |
12.37 |
|
|
|
| 21 |
A |
1267 |
1213 |
2.21 |
|
|
|
| 22 |
A |
1229 |
1176 |
5.01 |
|
|
|
| 23 |
A |
1202 |
1151 |
0.35 |
|
|
|
| 24 |
A |
1169 |
1118 |
4.30 |
|
|
|
| 25 |
A |
1127 |
1079 |
0.41 |
|
|
|
| 26 |
A |
1068 |
1022 |
0.76 |
|
|
|
| 27 |
A |
995 |
953 |
0.44 |
|
|
|
| 28 |
A |
982 |
940 |
0.09 |
|
|
|
| 29 |
A |
966 |
924 |
3.51 |
|
|
|
| 30 |
A |
944 |
904 |
0.15 |
|
|
|
| 31 |
A |
910 |
871 |
1.21 |
|
|
|
| 32 |
A |
876 |
838 |
4.64 |
|
|
|
| 33 |
A |
834 |
798 |
3.30 |
|
|
|
| 34 |
A |
809 |
774 |
1.79 |
|
|
|
| 35 |
A |
751 |
719 |
1.40 |
|
|
|
| 36 |
A |
616 |
589 |
0.45 |
|
|
|
| 37 |
A |
454 |
435 |
0.25 |
|
|
|
| 38 |
A |
363 |
347 |
1.00 |
|
|
|
| 39 |
A |
242 |
232 |
2.46 |
|
|
|
| 40 |
A |
187 |
179 |
14.57 |
|
|
|
| 41 |
A |
162 |
155 |
0.75 |
|
|
|
| 42 |
A |
40 |
38 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30835.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29512.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.116 |
|
|
|
| 2 |
H |
0.116 |
|
|
|
| 3 |
C |
-0.239 |
|
|
|
| 4 |
H |
0.116 |
|
|
|
| 5 |
H |
0.116 |
|
|
|
| 6 |
C |
-0.223 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
C |
-0.240 |
|
|
|
| 10 |
H |
0.117 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
C |
-0.211 |
|
|
|
| 13 |
H |
0.139 |
|
|
|
| 14 |
C |
-0.102 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
S |
-0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.510 |
0.690 |
0.618 |
1.771 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.326 |
2.456 |
1.116 |
| y |
2.456 |
-44.779 |
0.967 |
| z |
1.116 |
0.967 |
-45.607 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.133 |
2.456 |
1.116 |
| y |
2.456 |
2.187 |
0.967 |
| z |
1.116 |
0.967 |
0.946 |
|
| Polar |
|---|
| 3z2-r2 | 1.892 |
|---|
| x2-y2 | -3.546 |
|---|
| xy | 2.456 |
|---|
| xz | 1.116 |
|---|
| yz | 0.967 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.405 |
0.179 |
-0.090 |
| y |
0.179 |
10.649 |
0.192 |
| z |
-0.090 |
0.192 |
9.141 |
<r2> (average value of r
2) Å
2
| <r2> |
227.226 |
| (<r2>)1/2 |
15.074 |