Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -246.864344 |
| Energy at 298.15K | |
| HF Energy | -245.880243 |
| Nuclear repulsion energy | 157.014334 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.131 |
-0.327 |
0.000 |
| O2 |
-0.741 |
-0.609 |
0.000 |
| C3 |
0.000 |
0.585 |
0.000 |
| C4 |
1.436 |
0.256 |
0.000 |
| N5 |
2.575 |
0.029 |
0.000 |
| H6 |
-2.649 |
-1.283 |
0.000 |
| H7 |
-2.420 |
0.241 |
0.892 |
| H8 |
-2.420 |
0.241 |
-0.892 |
| H9 |
-0.216 |
1.194 |
-0.888 |
| H10 |
-0.216 |
1.194 |
0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4183 | 2.3184 | 3.6143 | 4.7199 | 1.0874 | 1.0964 | 1.0964 | 2.6016 | 2.6016 |
O2 | 1.4183 | | 1.4062 | 2.3427 | 3.3775 | 2.0232 | 2.0824 | 2.0824 | 2.0773 | 2.0773 | C3 | 2.3184 | 1.4062 | | 1.4730 | 2.6345 | 3.2418 | 2.6017 | 2.6017 | 1.0976 | 1.0976 | C4 | 3.6143 | 2.3427 | 1.4730 | | 1.1618 | 4.3650 | 3.9572 | 3.9572 | 2.0967 | 2.0967 | N5 | 4.7199 | 3.3775 | 2.6345 | 1.1618 | | 5.3865 | 5.0783 | 5.0783 | 3.1521 | 3.1521 | H6 | 1.0874 | 2.0232 | 3.2418 | 4.3650 | 5.3865 | | 1.7813 | 1.7813 | 3.5836 | 3.5836 | H7 | 1.0964 | 2.0824 | 2.6017 | 3.9572 | 5.0783 | 1.7813 | | 1.7840 | 2.9882 | 2.4004 | H8 | 1.0964 | 2.0824 | 2.6017 | 3.9572 | 5.0783 | 1.7813 | 1.7840 | | 2.4004 | 2.9882 | H9 | 2.6016 | 2.0773 | 1.0976 | 2.0967 | 3.1521 | 3.5836 | 2.9882 | 2.4004 | | 1.7753 | H10 | 2.6016 | 2.0773 | 1.0976 | 2.0967 | 3.1521 | 3.5836 | 2.4004 | 2.9882 | 1.7753 | |
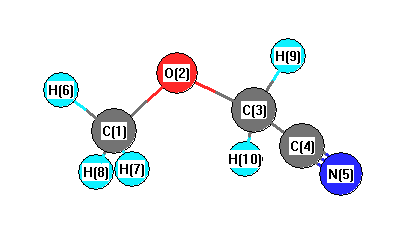 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.334 |
|
O2 |
C1 |
H6 |
106.947 |
| O2 |
C1 |
H7 |
111.161 |
|
O2 |
C1 |
H8 |
111.161 |
| O2 |
C3 |
C4 |
108.888 |
|
O2 |
C3 |
H9 |
111.535 |
| O2 |
C3 |
H10 |
111.535 |
|
C3 |
C4 |
N5 |
178.318 |
| C4 |
C3 |
H9 |
108.421 |
|
C4 |
C3 |
H10 |
108.421 |
| H6 |
C1 |
H7 |
109.316 |
|
H6 |
C1 |
H8 |
109.316 |
| H7 |
C1 |
H8 |
108.898 |
|
H9 |
C3 |
H10 |
107.945 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -246.866777 |
| Energy at 298.15K | |
| HF Energy | -245.881710 |
| Nuclear repulsion energy | 159.987990 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.542 |
-0.783 |
0.138 |
| O2 |
1.088 |
0.434 |
-0.441 |
| C3 |
-0.048 |
0.935 |
0.212 |
| C4 |
-1.238 |
0.063 |
0.035 |
| N5 |
-2.158 |
-0.635 |
-0.098 |
| H6 |
2.445 |
-1.066 |
-0.398 |
| H7 |
0.795 |
-1.576 |
0.034 |
| H8 |
1.776 |
-0.646 |
1.200 |
| H9 |
-0.268 |
1.911 |
-0.219 |
| H10 |
0.122 |
1.056 |
1.289 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4221 | 2.3420 | 2.9080 | 3.7107 | 1.0874 | 1.0946 | 1.0962 | 3.2651 | 2.5931 |
O2 | 1.4221 | | 1.4023 | 2.4030 | 3.4346 | 2.0235 | 2.0860 | 2.0815 | 2.0171 | 2.0764 | C3 | 2.3420 | 1.4023 | | 1.4857 | 2.6483 | 3.2543 | 2.6545 | 2.6083 | 1.0893 | 1.0972 | C4 | 2.9080 | 2.4030 | 1.4857 | | 1.1627 | 3.8769 | 2.6115 | 3.3082 | 2.1024 | 2.0994 | N5 | 3.7107 | 3.4346 | 2.6483 | 1.1627 | | 4.6332 | 3.1020 | 4.1427 | 3.1733 | 3.1597 | H6 | 1.0874 | 2.0235 | 3.2543 | 3.8769 | 4.6332 | | 1.7806 | 1.7822 | 4.0318 | 3.5700 | H7 | 1.0946 | 2.0860 | 2.6545 | 2.6115 | 3.1020 | 1.7806 | | 1.7851 | 3.6540 | 2.9926 | H8 | 1.0962 | 2.0815 | 2.6083 | 3.3082 | 4.1427 | 1.7822 | 1.7851 | | 3.5679 | 2.3752 | H9 | 3.2651 | 2.0171 | 1.0893 | 2.1024 | 3.1733 | 4.0318 | 3.6540 | 3.5679 | | 1.7765 | H10 | 2.5931 | 2.0764 | 1.0972 | 2.0994 | 3.1597 | 3.5700 | 2.9926 | 2.3752 | 1.7765 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.028 |
|
O2 |
C1 |
H6 |
106.722 |
| O2 |
C1 |
H7 |
111.297 |
|
O2 |
C1 |
H8 |
110.832 |
| O2 |
C3 |
C4 |
112.583 |
|
O2 |
C3 |
H9 |
107.436 |
| O2 |
C3 |
H10 |
111.764 |
|
C3 |
C4 |
N5 |
178.994 |
| C4 |
C3 |
H9 |
108.482 |
|
C4 |
C3 |
H10 |
107.795 |
| H6 |
C1 |
H7 |
109.382 |
|
H6 |
C1 |
H8 |
109.416 |
| H7 |
C1 |
H8 |
109.145 |
|
H9 |
C3 |
H10 |
108.676 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
