Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -246.873060 |
| Energy at 298.15K | -246.878660 |
| HF Energy | -245.879970 |
| Nuclear repulsion energy | 157.801349 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3204 |
3041 |
11.94 |
|
|
|
| 2 |
A' |
3053 |
2898 |
47.24 |
|
|
|
| 3 |
A' |
3046 |
2892 |
19.15 |
|
|
|
| 4 |
A' |
2235 |
2122 |
6.26 |
|
|
|
| 5 |
A' |
1544 |
1465 |
6.49 |
|
|
|
| 6 |
A' |
1525 |
1448 |
4.84 |
|
|
|
| 7 |
A' |
1496 |
1421 |
0.26 |
|
|
|
| 8 |
A' |
1427 |
1355 |
33.99 |
|
|
|
| 9 |
A' |
1241 |
1179 |
59.14 |
|
|
|
| 10 |
A' |
1185 |
1125 |
112.87 |
|
|
|
| 11 |
A' |
1020 |
969 |
30.33 |
|
|
|
| 12 |
A' |
969 |
920 |
7.05 |
|
|
|
| 13 |
A' |
546 |
518 |
1.12 |
|
|
|
| 14 |
A' |
380 |
360 |
1.90 |
|
|
|
| 15 |
A' |
179 |
170 |
2.32 |
|
|
|
| 16 |
A" |
3111 |
2953 |
35.28 |
|
|
|
| 17 |
A" |
3074 |
2918 |
21.46 |
|
|
|
| 18 |
A" |
1522 |
1445 |
8.55 |
|
|
|
| 19 |
A" |
1283 |
1219 |
3.42 |
|
|
|
| 20 |
A" |
1199 |
1138 |
3.74 |
|
|
|
| 21 |
A" |
1046 |
993 |
2.00 |
|
|
|
| 22 |
A" |
366 |
347 |
1.35 |
|
|
|
| 23 |
A" |
233 |
221 |
3.62 |
|
|
|
| 24 |
A" |
93 |
88 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17487.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 16602.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.118 |
-0.328 |
0.000 |
| O2 |
-0.736 |
-0.611 |
0.000 |
| C3 |
0.000 |
0.580 |
0.000 |
| C4 |
1.421 |
0.259 |
0.000 |
| N5 |
2.564 |
0.036 |
0.000 |
| H6 |
-2.636 |
-1.277 |
0.000 |
| H7 |
-2.404 |
0.238 |
0.887 |
| H8 |
-2.404 |
0.238 |
-0.887 |
| H9 |
-0.220 |
1.185 |
-0.882 |
| H10 |
-0.220 |
1.185 |
0.882 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4110 | 2.3041 | 3.5866 | 4.6963 | 1.0812 | 1.0901 | 1.0901 | 2.5825 | 2.5825 |
O2 | 1.4110 | | 1.3999 | 2.3250 | 3.3627 | 2.0134 | 2.0715 | 2.0715 | 2.0666 | 2.0666 | C3 | 2.3041 | 1.3999 | | 1.4563 | 2.6216 | 3.2241 | 2.5850 | 2.5850 | 1.0921 | 1.0921 | C4 | 3.5866 | 2.3250 | 1.4563 | | 1.1656 | 4.3372 | 3.9260 | 3.9260 | 2.0801 | 2.0801 | N5 | 4.6963 | 3.3627 | 2.6216 | 1.1656 | | 5.3632 | 5.0510 | 5.0510 | 3.1389 | 3.1389 | H6 | 1.0812 | 2.0134 | 3.2241 | 4.3372 | 5.3632 | | 1.7706 | 1.7706 | 3.5603 | 3.5603 | H7 | 1.0901 | 2.0715 | 2.5850 | 3.9260 | 5.0510 | 1.7706 | | 1.7732 | 2.9657 | 2.3806 | H8 | 1.0901 | 2.0715 | 2.5850 | 3.9260 | 5.0510 | 1.7706 | 1.7732 | | 2.3806 | 2.9657 | H9 | 2.5825 | 2.0666 | 1.0921 | 2.0801 | 3.1389 | 3.5603 | 2.9657 | 2.3806 | | 1.7641 | H10 | 2.5825 | 2.0666 | 1.0921 | 2.0801 | 3.1389 | 3.5603 | 2.3806 | 2.9657 | 1.7641 | |
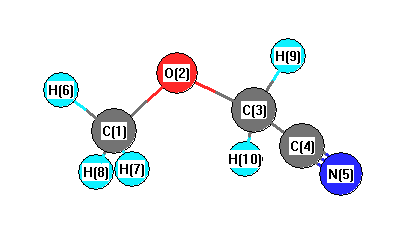 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.110 |
|
O2 |
C1 |
H6 |
107.037 |
| O2 |
C1 |
H7 |
111.191 |
|
O2 |
C1 |
H8 |
111.191 |
| O2 |
C3 |
C4 |
108.965 |
|
O2 |
C3 |
H9 |
111.456 |
| O2 |
C3 |
H10 |
111.456 |
|
C3 |
C4 |
N5 |
178.321 |
| C4 |
C3 |
H9 |
108.576 |
|
C4 |
C3 |
H10 |
108.576 |
| H6 |
C1 |
H7 |
109.265 |
|
H6 |
C1 |
H8 |
109.265 |
| H7 |
C1 |
H8 |
108.847 |
|
H9 |
C3 |
H10 |
107.728 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -246.875530 |
| Energy at 298.15K | -246.881229 |
| HF Energy | -245.881312 |
| Nuclear repulsion energy | 160.901380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3204 |
3042 |
12.48 |
|
|
|
| 2 |
A |
3165 |
3005 |
2.91 |
|
|
|
| 3 |
A |
3126 |
2967 |
25.89 |
|
|
|
| 4 |
A |
3061 |
2906 |
34.30 |
|
|
|
| 5 |
A |
3059 |
2904 |
23.60 |
|
|
|
| 6 |
A |
2217 |
2105 |
6.47 |
|
|
|
| 7 |
A |
1541 |
1463 |
9.20 |
|
|
|
| 8 |
A |
1524 |
1447 |
8.12 |
|
|
|
| 9 |
A |
1505 |
1429 |
4.37 |
|
|
|
| 10 |
A |
1495 |
1419 |
0.85 |
|
|
|
| 11 |
A |
1405 |
1334 |
14.21 |
|
|
|
| 12 |
A |
1331 |
1264 |
6.65 |
|
|
|
| 13 |
A |
1236 |
1173 |
48.90 |
|
|
|
| 14 |
A |
1200 |
1139 |
9.91 |
|
|
|
| 15 |
A |
1180 |
1121 |
88.36 |
|
|
|
| 16 |
A |
1040 |
987 |
7.62 |
|
|
|
| 17 |
A |
970 |
921 |
29.70 |
|
|
|
| 18 |
A |
920 |
874 |
14.00 |
|
|
|
| 19 |
A |
602 |
571 |
1.91 |
|
|
|
| 20 |
A |
394 |
374 |
2.81 |
|
|
|
| 21 |
A |
356 |
338 |
1.03 |
|
|
|
| 22 |
A |
260 |
247 |
9.69 |
|
|
|
| 23 |
A |
179 |
170 |
3.94 |
|
|
|
| 24 |
A |
117 |
111 |
8.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17541.7 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 16654.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.536 |
-0.770 |
0.138 |
| O2 |
1.072 |
0.425 |
-0.445 |
| C3 |
-0.045 |
0.925 |
0.211 |
| C4 |
-1.228 |
0.065 |
0.040 |
| N5 |
-2.147 |
-0.631 |
-0.099 |
| H6 |
2.426 |
-1.057 |
-0.405 |
| H7 |
0.797 |
-1.566 |
0.060 |
| H8 |
1.788 |
-0.623 |
1.189 |
| H9 |
-0.266 |
1.900 |
-0.208 |
| H10 |
0.124 |
1.037 |
1.284 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4091 | 2.3195 | 2.8892 | 3.6938 | 1.0811 | 1.0893 | 1.0902 | 3.2399 | 2.5642 |
O2 | 1.4091 | | 1.3882 | 2.3780 | 3.4055 | 2.0081 | 2.0730 | 2.0690 | 2.0050 | 2.0646 | C3 | 2.3195 | 1.3882 | | 1.4730 | 2.6339 | 3.2270 | 2.6342 | 2.5909 | 1.0834 | 1.0918 | C4 | 2.8892 | 2.3780 | 1.4730 | | 1.1610 | 3.8481 | 2.6001 | 3.2998 | 2.0866 | 2.0781 | N5 | 3.6938 | 3.4055 | 2.6339 | 1.1610 | | 4.6033 | 3.0933 | 4.1408 | 3.1549 | 3.1385 | H6 | 1.0811 | 2.0081 | 3.2270 | 3.8481 | 4.6033 | | 1.7691 | 1.7704 | 4.0035 | 3.5410 | H7 | 1.0893 | 2.0730 | 2.6342 | 2.6001 | 3.0933 | 1.7691 | | 1.7740 | 3.6352 | 2.9546 | H8 | 1.0902 | 2.0690 | 2.5909 | 3.2998 | 4.1408 | 1.7704 | 1.7740 | | 3.5402 | 2.3529 | H9 | 3.2399 | 2.0050 | 1.0834 | 2.0866 | 3.1549 | 4.0035 | 3.6352 | 3.5402 | | 1.7664 | H10 | 2.5642 | 2.0646 | 1.0918 | 2.0781 | 3.1385 | 3.5410 | 2.9546 | 2.3529 | 1.7664 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.028 |
|
O2 |
C1 |
H6 |
106.753 |
| O2 |
C1 |
H7 |
111.506 |
|
O2 |
C1 |
H8 |
111.117 |
| O2 |
C3 |
C4 |
112.395 |
|
O2 |
C3 |
H9 |
107.795 |
| O2 |
C3 |
H10 |
112.165 |
|
C3 |
C4 |
N5 |
178.908 |
| C4 |
C3 |
H9 |
108.464 |
|
C4 |
C3 |
H10 |
107.312 |
| H6 |
C1 |
H7 |
109.194 |
|
H6 |
C1 |
H8 |
109.248 |
| H7 |
C1 |
H8 |
108.966 |
|
H9 |
C3 |
H10 |
108.599 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
