Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -247.240007 |
| Energy at 298.15K | -247.245644 |
| HF Energy | -247.240007 |
| Nuclear repulsion energy | 161.125997 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3029 |
15.87 |
|
|
|
| 2 |
A |
3123 |
2989 |
3.56 |
|
|
|
| 3 |
A |
3080 |
2948 |
31.58 |
|
|
|
| 4 |
A |
3017 |
2887 |
41.19 |
|
|
|
| 5 |
A |
3014 |
2885 |
25.83 |
|
|
|
| 6 |
A |
2385 |
2283 |
0.20 |
|
|
|
| 7 |
A |
1508 |
1443 |
9.81 |
|
|
|
| 8 |
A |
1488 |
1424 |
9.69 |
|
|
|
| 9 |
A |
1478 |
1414 |
0.80 |
|
|
|
| 10 |
A |
1468 |
1405 |
4.44 |
|
|
|
| 11 |
A |
1388 |
1329 |
13.59 |
|
|
|
| 12 |
A |
1314 |
1258 |
8.99 |
|
|
|
| 13 |
A |
1226 |
1173 |
73.77 |
|
|
|
| 14 |
A |
1181 |
1131 |
16.55 |
|
|
|
| 15 |
A |
1175 |
1125 |
67.92 |
|
|
|
| 16 |
A |
1031 |
987 |
8.28 |
|
|
|
| 17 |
A |
968 |
926 |
29.40 |
|
|
|
| 18 |
A |
911 |
872 |
16.28 |
|
|
|
| 19 |
A |
607 |
581 |
1.63 |
|
|
|
| 20 |
A |
380 |
364 |
1.61 |
|
|
|
| 21 |
A |
361 |
346 |
2.58 |
|
|
|
| 22 |
A |
247 |
236 |
10.66 |
|
|
|
| 23 |
A |
167 |
160 |
2.63 |
|
|
|
| 24 |
A |
112 |
108 |
8.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17396.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 16650.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.546 |
-0.769 |
0.134 |
| O2 |
1.076 |
0.435 |
-0.429 |
| C3 |
-0.061 |
0.925 |
0.205 |
| C4 |
-1.233 |
0.054 |
0.034 |
| N5 |
-2.140 |
-0.636 |
-0.096 |
| H6 |
2.443 |
-1.042 |
-0.412 |
| H7 |
0.808 |
-1.568 |
0.037 |
| H8 |
1.793 |
-0.638 |
1.191 |
| H9 |
-0.286 |
1.896 |
-0.231 |
| H10 |
0.099 |
1.064 |
1.280 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4099 | 2.3369 | 2.9005 | 3.6956 | 1.0846 | 1.0919 | 1.0935 | 3.2549 | 2.6022 |
O2 | 1.4099 | | 1.3916 | 2.3858 | 3.4062 | 2.0127 | 2.0744 | 2.0712 | 2.0073 | 2.0669 | C3 | 2.3369 | 1.3916 | | 1.4699 | 2.6171 | 3.2440 | 2.6463 | 2.6186 | 1.0876 | 1.0957 | C4 | 2.9005 | 2.3858 | 1.4699 | | 1.1473 | 3.8618 | 2.6078 | 3.3132 | 2.0874 | 2.0844 | N5 | 3.6956 | 3.4062 | 2.6171 | 1.1473 | | 4.6114 | 3.0949 | 4.1382 | 3.1410 | 3.1295 | H6 | 1.0846 | 2.0127 | 3.2440 | 3.8618 | 4.6114 | | 1.7748 | 1.7760 | 4.0142 | 3.5772 | H7 | 1.0919 | 2.0744 | 2.6463 | 2.6078 | 3.0949 | 1.7748 | | 1.7791 | 3.6429 | 2.9962 | H8 | 1.0935 | 2.0712 | 2.6186 | 3.3132 | 4.1382 | 1.7760 | 1.7791 | | 3.5735 | 2.4039 | H9 | 3.2549 | 2.0073 | 1.0876 | 2.0874 | 3.1410 | 4.0142 | 3.6429 | 3.5735 | | 1.7677 | H10 | 2.6022 | 2.0669 | 1.0957 | 2.0844 | 3.1295 | 3.5772 | 2.9962 | 2.4039 | 1.7677 | |
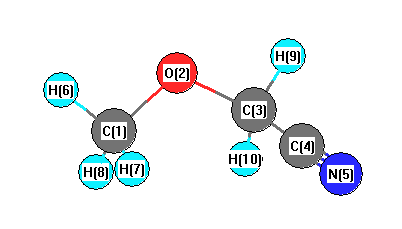 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.059 |
|
O2 |
C1 |
H6 |
106.854 |
| O2 |
C1 |
H7 |
111.391 |
|
O2 |
C1 |
H8 |
111.029 |
| O2 |
C3 |
C4 |
112.952 |
|
O2 |
C3 |
H9 |
107.491 |
| O2 |
C3 |
H10 |
111.857 |
|
C3 |
C4 |
N5 |
179.300 |
| C4 |
C3 |
H9 |
108.488 |
|
C4 |
C3 |
H10 |
107.786 |
| H6 |
C1 |
H7 |
109.264 |
|
H6 |
C1 |
H8 |
109.260 |
| H7 |
C1 |
H8 |
108.994 |
|
H9 |
C3 |
H10 |
108.122 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.156 |
|
|
|
| 2 |
O |
-0.245 |
|
|
|
| 3 |
C |
-0.021 |
|
|
|
| 4 |
C |
-0.107 |
|
|
|
| 5 |
N |
-0.050 |
|
|
|
| 6 |
H |
0.130 |
|
|
|
| 7 |
H |
0.101 |
|
|
|
| 8 |
H |
0.083 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.573 |
1.288 |
1.337 |
3.172 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.130 |
-5.432 |
0.553 |
| y |
-5.432 |
-27.910 |
0.581 |
| z |
0.553 |
0.581 |
-29.629 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.361 |
-5.432 |
0.553 |
| y |
-5.432 |
4.969 |
0.581 |
| z |
0.553 |
0.581 |
2.391 |
|
| Polar |
|---|
| 3z2-r2 | 4.783 |
|---|
| x2-y2 | -8.220 |
|---|
| xy | -5.432 |
|---|
| xz | 0.553 |
|---|
| yz | 0.581 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.487 |
0.356 |
0.168 |
| y |
0.356 |
6.159 |
0.184 |
| z |
0.168 |
0.184 |
5.077 |
<r2> (average value of r
2) Å
2
| <r2> |
121.158 |
| (<r2>)1/2 |
11.007 |
