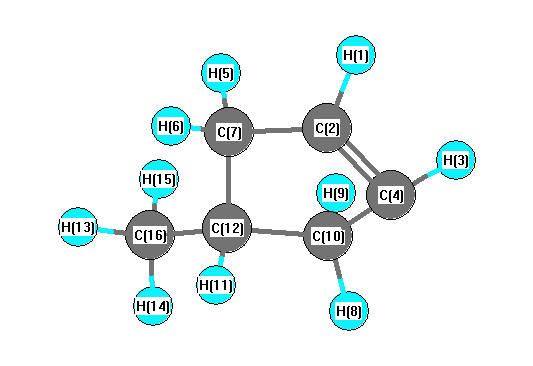
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3221 |
3083 |
27.41 |
|
|
|
| 2 |
A |
3195 |
3058 |
8.09 |
|
|
|
| 3 |
A |
3120 |
2986 |
26.16 |
|
|
|
| 4 |
A |
3117 |
2983 |
48.71 |
|
|
|
| 5 |
A |
3086 |
2953 |
26.41 |
|
|
|
| 6 |
A |
3081 |
2948 |
37.19 |
|
|
|
| 7 |
A |
3048 |
2917 |
33.98 |
|
|
|
| 8 |
A |
3045 |
2914 |
13.70 |
|
|
|
| 9 |
A |
3015 |
2885 |
40.52 |
|
|
|
| 10 |
A |
3013 |
2884 |
24.57 |
|
|
|
| 11 |
A |
1705 |
1632 |
3.87 |
|
|
|
| 12 |
A |
1500 |
1436 |
7.74 |
|
|
|
| 13 |
A |
1495 |
1431 |
3.02 |
|
|
|
| 14 |
A |
1490 |
1426 |
0.68 |
|
|
|
| 15 |
A |
1482 |
1418 |
2.55 |
|
|
|
| 16 |
A |
1407 |
1347 |
4.59 |
|
|
|
| 17 |
A |
1383 |
1324 |
0.78 |
|
|
|
| 18 |
A |
1377 |
1318 |
4.43 |
|
|
|
| 19 |
A |
1338 |
1280 |
1.60 |
|
|
|
| 20 |
A |
1308 |
1252 |
0.36 |
|
|
|
| 21 |
A |
1295 |
1240 |
3.62 |
|
|
|
| 22 |
A |
1240 |
1186 |
0.25 |
|
|
|
| 23 |
A |
1177 |
1127 |
0.00 |
|
|
|
| 24 |
A |
1154 |
1104 |
2.48 |
|
|
|
| 25 |
A |
1135 |
1086 |
0.45 |
|
|
|
| 26 |
A |
1116 |
1068 |
0.60 |
|
|
|
| 27 |
A |
1076 |
1029 |
3.18 |
|
|
|
| 28 |
A |
997 |
955 |
10.68 |
|
|
|
| 29 |
A |
996 |
954 |
0.69 |
|
|
|
| 30 |
A |
985 |
943 |
0.16 |
|
|
|
| 31 |
A |
932 |
892 |
4.52 |
|
|
|
| 32 |
A |
929 |
889 |
5.89 |
|
|
|
| 33 |
A |
905 |
867 |
0.01 |
|
|
|
| 34 |
A |
838 |
802 |
1.41 |
|
|
|
| 35 |
A |
782 |
748 |
0.52 |
|
|
|
| 36 |
A |
709 |
678 |
36.11 |
|
|
|
| 37 |
A |
569 |
545 |
3.23 |
|
|
|
| 38 |
A |
412 |
394 |
0.88 |
|
|
|
| 39 |
A |
400 |
382 |
0.08 |
|
|
|
| 40 |
A |
306 |
292 |
0.03 |
|
|
|
| 41 |
A |
246 |
235 |
0.00 |
|
|
|
| 42 |
A |
105 |
100 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31863.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30496.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.135 |
|
|
|
| 2 |
C |
-0.173 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
C |
-0.173 |
|
|
|
| 5 |
H |
0.103 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
C |
-0.183 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.103 |
|
|
|
| 10 |
C |
-0.183 |
|
|
|
| 11 |
H |
0.121 |
|
|
|
| 12 |
C |
-0.071 |
|
|
|
| 13 |
H |
0.118 |
|
|
|
| 14 |
H |
0.118 |
|
|
|
| 15 |
H |
0.112 |
|
|
|
| 16 |
C |
-0.382 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.213 |
0.000 |
0.026 |
0.215 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.354 |
0.000 |
-0.417 |
| y |
0.000 |
-37.097 |
0.000 |
| z |
-0.417 |
0.000 |
-39.115 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.248 |
0.000 |
-0.417 |
| y |
0.000 |
1.638 |
0.000 |
| z |
-0.417 |
0.000 |
-1.390 |
|
| Polar |
|---|
| 3z2-r2 | -2.779 |
|---|
| x2-y2 | -1.257 |
|---|
| xy | 0.000 |
|---|
| xz | -0.417 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.834 |
0.000 |
-0.005 |
| y |
0.000 |
10.969 |
0.000 |
| z |
-0.005 |
0.000 |
7.839 |
<r2> (average value of r
2) Å
2
| <r2> |
162.267 |
| (<r2>)1/2 |
12.738 |