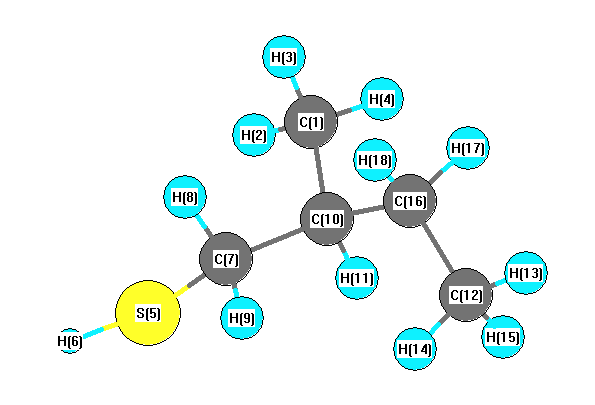
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3143 |
3008 |
15.61 |
|
|
|
| 2 |
A |
3132 |
2998 |
34.01 |
|
|
|
| 3 |
A |
3128 |
2994 |
31.84 |
|
|
|
| 4 |
A |
3124 |
2990 |
48.52 |
|
|
|
| 5 |
A |
3119 |
2985 |
17.15 |
|
|
|
| 6 |
A |
3076 |
2944 |
18.75 |
|
|
|
| 7 |
A |
3069 |
2938 |
22.75 |
|
|
|
| 8 |
A |
3055 |
2924 |
26.42 |
|
|
|
| 9 |
A |
3049 |
2918 |
33.06 |
|
|
|
| 10 |
A |
3040 |
2910 |
15.36 |
|
|
|
| 11 |
A |
3031 |
2901 |
3.53 |
|
|
|
| 12 |
A |
2718 |
2601 |
4.37 |
|
|
|
| 13 |
A |
1508 |
1444 |
8.86 |
|
|
|
| 14 |
A |
1499 |
1435 |
12.33 |
|
|
|
| 15 |
A |
1499 |
1435 |
3.30 |
|
|
|
| 16 |
A |
1496 |
1432 |
1.41 |
|
|
|
| 17 |
A |
1483 |
1420 |
1.91 |
|
|
|
| 18 |
A |
1475 |
1412 |
5.76 |
|
|
|
| 19 |
A |
1412 |
1352 |
4.07 |
|
|
|
| 20 |
A |
1412 |
1351 |
10.08 |
|
|
|
| 21 |
A |
1387 |
1327 |
2.27 |
|
|
|
| 22 |
A |
1372 |
1313 |
0.65 |
|
|
|
| 23 |
A |
1325 |
1268 |
0.91 |
|
|
|
| 24 |
A |
1302 |
1246 |
7.95 |
|
|
|
| 25 |
A |
1273 |
1218 |
3.69 |
|
|
|
| 26 |
A |
1228 |
1175 |
7.25 |
|
|
|
| 27 |
A |
1187 |
1136 |
0.55 |
|
|
|
| 28 |
A |
1151 |
1101 |
2.60 |
|
|
|
| 29 |
A |
1091 |
1044 |
1.01 |
|
|
|
| 30 |
A |
1055 |
1010 |
0.64 |
|
|
|
| 31 |
A |
1031 |
987 |
2.15 |
|
|
|
| 32 |
A |
977 |
935 |
5.05 |
|
|
|
| 33 |
A |
950 |
909 |
1.08 |
|
|
|
| 34 |
A |
900 |
861 |
2.74 |
|
|
|
| 35 |
A |
868 |
831 |
0.91 |
|
|
|
| 36 |
A |
799 |
765 |
1.58 |
|
|
|
| 37 |
A |
792 |
758 |
6.63 |
|
|
|
| 38 |
A |
729 |
698 |
0.79 |
|
|
|
| 39 |
A |
447 |
428 |
0.11 |
|
|
|
| 40 |
A |
394 |
377 |
0.09 |
|
|
|
| 41 |
A |
371 |
355 |
0.88 |
|
|
|
| 42 |
A |
254 |
243 |
0.33 |
|
|
|
| 43 |
A |
242 |
231 |
0.06 |
|
|
|
| 44 |
A |
206 |
197 |
0.53 |
|
|
|
| 45 |
A |
198 |
190 |
1.33 |
|
|
|
| 46 |
A |
170 |
162 |
13.10 |
|
|
|
| 47 |
A |
95 |
91 |
0.73 |
|
|
|
| 48 |
A |
67 |
64 |
4.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35163.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 33655.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.364 |
|
|
|
| 2 |
H |
0.134 |
|
|
|
| 3 |
H |
0.116 |
|
|
|
| 4 |
H |
0.115 |
|
|
|
| 5 |
S |
-0.199 |
|
|
|
| 6 |
H |
0.108 |
|
|
|
| 7 |
C |
-0.222 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
C |
-0.106 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
C |
-0.376 |
|
|
|
| 13 |
H |
0.123 |
|
|
|
| 14 |
H |
0.115 |
|
|
|
| 15 |
H |
0.119 |
|
|
|
| 16 |
C |
-0.214 |
|
|
|
| 17 |
H |
0.117 |
|
|
|
| 18 |
H |
0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.408 |
-0.587 |
0.655 |
1.660 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.666 |
2.140 |
-1.274 |
| y |
2.140 |
-46.057 |
-1.521 |
| z |
-1.274 |
-1.521 |
-47.939 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.668 |
2.140 |
-1.274 |
| y |
2.140 |
2.745 |
-1.521 |
| z |
-1.274 |
-1.521 |
-0.077 |
|
| Polar |
|---|
| 3z2-r2 | -0.155 |
|---|
| x2-y2 | -3.608 |
|---|
| xy | 2.140 |
|---|
| xz | -1.274 |
|---|
| yz | -1.521 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.974 |
0.259 |
0.208 |
| y |
0.259 |
11.553 |
-0.194 |
| z |
0.208 |
-0.194 |
9.790 |
<r2> (average value of r
2) Å
2
| <r2> |
282.963 |
| (<r2>)1/2 |
16.822 |