Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.392764 |
| Energy at 298.15K | -235.406361 |
| Nuclear repulsion energy | 257.451934 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3177 |
3017 |
42.00 |
|
|
|
| 2 |
A' |
3150 |
2991 |
34.50 |
|
|
|
| 3 |
A' |
3148 |
2989 |
34.19 |
|
|
|
| 4 |
A' |
3144 |
2985 |
6.93 |
|
|
|
| 5 |
A' |
3123 |
2965 |
22.36 |
|
|
|
| 6 |
A' |
3098 |
2941 |
7.21 |
|
|
|
| 7 |
A' |
3072 |
2916 |
26.35 |
|
|
|
| 8 |
A' |
3067 |
2912 |
24.77 |
|
|
|
| 9 |
A' |
1530 |
1453 |
3.41 |
|
|
|
| 10 |
A' |
1527 |
1450 |
1.55 |
|
|
|
| 11 |
A' |
1514 |
1438 |
6.88 |
|
|
|
| 12 |
A' |
1500 |
1424 |
0.23 |
|
|
|
| 13 |
A' |
1418 |
1346 |
3.18 |
|
|
|
| 14 |
A' |
1407 |
1336 |
13.63 |
|
|
|
| 15 |
A' |
1335 |
1267 |
2.50 |
|
|
|
| 16 |
A' |
1320 |
1253 |
6.89 |
|
|
|
| 17 |
A' |
1247 |
1184 |
0.38 |
|
|
|
| 18 |
A' |
1220 |
1158 |
0.21 |
|
|
|
| 19 |
A' |
1032 |
980 |
0.56 |
|
|
|
| 20 |
A' |
1012 |
961 |
0.67 |
|
|
|
| 21 |
A' |
964 |
915 |
0.29 |
|
|
|
| 22 |
A' |
944 |
896 |
0.47 |
|
|
|
| 23 |
A' |
894 |
848 |
0.26 |
|
|
|
| 24 |
A' |
720 |
684 |
0.80 |
|
|
|
| 25 |
A' |
607 |
576 |
1.35 |
|
|
|
| 26 |
A' |
391 |
371 |
0.48 |
|
|
|
| 27 |
A' |
353 |
336 |
0.01 |
|
|
|
| 28 |
A' |
192 |
182 |
0.00 |
|
|
|
| 29 |
A" |
3149 |
2990 |
19.28 |
|
|
|
| 30 |
A" |
3143 |
2984 |
27.08 |
|
|
|
| 31 |
A" |
3138 |
2979 |
11.10 |
|
|
|
| 32 |
A" |
3095 |
2939 |
58.94 |
|
|
|
| 33 |
A" |
1528 |
1450 |
8.57 |
|
|
|
| 34 |
A" |
1505 |
1429 |
0.12 |
|
|
|
| 35 |
A" |
1492 |
1416 |
4.73 |
|
|
|
| 36 |
A" |
1284 |
1219 |
0.52 |
|
|
|
| 37 |
A" |
1266 |
1202 |
0.12 |
|
|
|
| 38 |
A" |
1236 |
1174 |
0.00 |
|
|
|
| 39 |
A" |
1193 |
1133 |
2.07 |
|
|
|
| 40 |
A" |
1096 |
1041 |
0.07 |
|
|
|
| 41 |
A" |
995 |
945 |
0.02 |
|
|
|
| 42 |
A" |
961 |
912 |
2.06 |
|
|
|
| 43 |
A" |
900 |
855 |
0.00 |
|
|
|
| 44 |
A" |
802 |
762 |
0.12 |
|
|
|
| 45 |
A" |
381 |
362 |
0.01 |
|
|
|
| 46 |
A" |
301 |
286 |
0.01 |
|
|
|
| 47 |
A" |
280 |
265 |
0.00 |
|
|
|
| 48 |
A" |
232 |
220 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37040.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 35166.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.112 |
0.400 |
0.000 |
| C2 |
0.406 |
-1.693 |
0.000 |
| C3 |
-0.112 |
-0.714 |
1.069 |
| C4 |
-0.112 |
-0.714 |
-1.069 |
| C5 |
1.202 |
1.153 |
0.000 |
| C6 |
-1.284 |
1.348 |
0.000 |
| H7 |
1.488 |
-1.756 |
0.000 |
| H8 |
-0.002 |
-2.696 |
0.000 |
| H9 |
0.491 |
-0.554 |
1.958 |
| H10 |
0.491 |
-0.554 |
-1.958 |
| H11 |
-1.127 |
-0.964 |
1.365 |
| H12 |
-1.127 |
-0.964 |
-1.365 |
| H13 |
2.047 |
0.470 |
0.000 |
| H14 |
-2.222 |
0.800 |
0.000 |
| H15 |
1.281 |
1.786 |
-0.881 |
| H16 |
1.281 |
1.786 |
0.881 |
| H17 |
-1.265 |
1.988 |
0.880 |
| H18 |
-1.265 |
1.988 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1562 | 1.5444 | 1.5444 | 1.5137 | 1.5072 | 2.6846 | 3.0983 | 2.2601 | 2.2601 | 2.1802 | 2.1802 | 2.1595 | 2.1483 | 2.1533 | 2.1533 | 2.1506 | 2.1506 |
C2 | 2.1562 | | 1.5393 | 1.5393 | 2.9555 | 3.4785 | 1.0839 | 1.0830 | 2.2668 | 2.2668 | 2.1779 | 2.1779 | 2.7152 | 3.6222 | 3.6944 | 3.6944 | 4.1368 | 4.1368 | C3 | 1.5444 | 1.5393 | | 2.1387 | 2.5209 | 2.6017 | 2.1879 | 2.2548 | 1.0858 | 3.0910 | 1.0864 | 2.6491 | 2.6841 | 2.8091 | 3.4634 | 2.8682 | 2.9439 | 3.5257 | C4 | 1.5444 | 1.5393 | 2.1387 | | 2.5209 | 2.6017 | 2.1879 | 2.2548 | 3.0910 | 1.0858 | 2.6491 | 1.0864 | 2.6841 | 2.8091 | 2.8682 | 3.4634 | 3.5257 | 2.9439 | C5 | 1.5137 | 2.9555 | 2.5209 | 2.5209 | | 2.4929 | 2.9234 | 4.0333 | 2.6931 | 2.6931 | 3.4301 | 3.4301 | 1.0868 | 3.4422 | 1.0878 | 1.0878 | 2.7485 | 2.7485 | C6 | 1.5072 | 3.4785 | 2.6017 | 2.6017 | 2.4929 | | 4.1611 | 4.2421 | 3.2561 | 3.2561 | 2.6888 | 2.6888 | 3.4442 | 1.0870 | 2.7469 | 2.7469 | 1.0883 | 1.0883 | H7 | 2.6846 | 1.0839 | 2.1879 | 2.1879 | 2.9234 | 4.1611 | | 1.7618 | 2.5042 | 2.5042 | 3.0539 | 3.0539 | 2.2954 | 4.5052 | 3.6562 | 3.6562 | 4.7295 | 4.7295 | H8 | 3.0983 | 1.0830 | 2.2548 | 2.2548 | 4.0333 | 4.2421 | 1.7618 | | 2.9438 | 2.9438 | 2.4756 | 2.4756 | 3.7715 | 4.1413 | 4.7451 | 4.7451 | 4.9304 | 4.9304 | H9 | 2.2601 | 2.2668 | 1.0858 | 3.0910 | 2.6931 | 3.2561 | 2.5042 | 2.9438 | | 3.9159 | 1.7717 | 3.7185 | 2.7021 | 3.6099 | 3.7631 | 2.6946 | 3.2723 | 4.1952 | H10 | 2.2601 | 2.2668 | 3.0910 | 1.0858 | 2.6931 | 3.2561 | 2.5042 | 2.9438 | 3.9159 | | 3.7185 | 1.7717 | 2.7021 | 3.6099 | 2.6946 | 3.7631 | 4.1952 | 3.2723 | H11 | 2.1802 | 2.1779 | 1.0864 | 2.6491 | 3.4301 | 2.6888 | 3.0539 | 2.4756 | 1.7717 | 3.7185 | | 2.7295 | 3.7403 | 2.4844 | 4.2899 | 3.6870 | 2.9942 | 3.7106 | H12 | 2.1802 | 2.1779 | 2.6491 | 1.0864 | 3.4301 | 2.6888 | 3.0539 | 2.4756 | 3.7185 | 1.7717 | 2.7295 | | 3.7403 | 2.4844 | 3.6870 | 4.2899 | 3.7106 | 2.9942 | H13 | 2.1595 | 2.7152 | 2.6841 | 2.6841 | 1.0868 | 3.4442 | 2.2954 | 3.7715 | 2.7021 | 2.7021 | 3.7403 | 3.7403 | | 4.2819 | 1.7593 | 1.7593 | 3.7476 | 3.7476 | H14 | 2.1483 | 3.6222 | 2.8091 | 2.8091 | 3.4422 | 1.0870 | 4.5052 | 4.1413 | 3.6099 | 3.6099 | 2.4844 | 2.4844 | 4.2819 | | 3.7447 | 3.7447 | 1.7615 | 1.7615 | H15 | 2.1533 | 3.6944 | 3.4634 | 2.8682 | 1.0878 | 2.7469 | 3.6562 | 4.7451 | 3.7631 | 2.6946 | 4.2899 | 3.6870 | 1.7593 | 3.7447 | | 1.7620 | 3.1020 | 2.5537 | H16 | 2.1533 | 3.6944 | 2.8682 | 3.4634 | 1.0878 | 2.7469 | 3.6562 | 4.7451 | 2.6946 | 3.7631 | 3.6870 | 4.2899 | 1.7593 | 3.7447 | 1.7620 | | 2.5537 | 3.1020 | H17 | 2.1506 | 4.1368 | 2.9439 | 3.5257 | 2.7485 | 1.0883 | 4.7295 | 4.9304 | 3.2723 | 4.1952 | 2.9942 | 3.7106 | 3.7476 | 1.7615 | 3.1020 | 2.5537 | | 1.7599 | H18 | 2.1506 | 4.1368 | 3.5257 | 2.9439 | 2.7485 | 1.0883 | 4.7295 | 4.9304 | 4.1952 | 3.2723 | 3.7106 | 2.9942 | 3.7476 | 1.7615 | 2.5537 | 3.1020 | 1.7599 | |
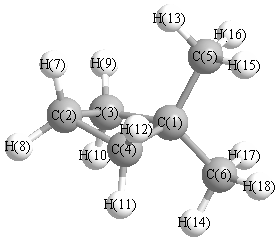 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.729 |
|
C1 |
C3 |
H9 |
117.409 |
| C1 |
C3 |
H11 |
110.729 |
|
C1 |
C4 |
C2 |
88.729 |
| C1 |
C4 |
H10 |
117.409 |
|
C1 |
C4 |
H12 |
110.729 |
| C1 |
C5 |
H13 |
111.221 |
|
C1 |
C5 |
H15 |
110.658 |
| C1 |
C5 |
H16 |
110.658 |
|
C1 |
C6 |
H14 |
110.769 |
| C1 |
C6 |
H17 |
110.871 |
|
C1 |
C6 |
H18 |
110.871 |
| C2 |
C3 |
H9 |
118.404 |
|
C2 |
C3 |
H11 |
110.904 |
| C2 |
C4 |
H10 |
118.404 |
|
C2 |
C4 |
H12 |
110.904 |
| C3 |
C1 |
C4 |
87.642 |
|
C3 |
C1 |
C5 |
111.034 |
| C3 |
C1 |
C6 |
116.975 |
|
C3 |
C2 |
C4 |
88.011 |
| C3 |
C2 |
H7 |
111.867 |
|
C3 |
C2 |
H8 |
117.551 |
| C4 |
C1 |
C5 |
111.034 |
|
C4 |
C1 |
C6 |
116.975 |
| C4 |
C2 |
H7 |
111.867 |
|
C4 |
C2 |
H8 |
117.551 |
| C5 |
C1 |
C6 |
111.215 |
|
H7 |
C2 |
H8 |
108.793 |
| H9 |
C3 |
H11 |
109.293 |
|
H10 |
C4 |
H12 |
109.293 |
| H13 |
C5 |
H15 |
108.008 |
|
H13 |
C5 |
H16 |
108.008 |
| H14 |
C6 |
H17 |
108.150 |
|
H14 |
C6 |
H18 |
108.150 |
| H15 |
C5 |
H16 |
108.171 |
|
H17 |
C6 |
H18 |
107.914 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
