Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULLultrafine/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.345477 |
| Energy at 298.15K | |
| HF Energy | -1592.072691 |
| Nuclear repulsion energy | 348.922360 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
517 |
494 |
0.00 |
40.74 |
0.04 |
0.08 |
| 2 |
A1 |
201 |
192 |
0.00 |
2.52 |
0.75 |
0.86 |
| 3 |
B1 |
456 |
436 |
0.00 |
18.00 |
0.75 |
0.86 |
| 4 |
B2 |
301 |
288 |
0.00 |
8.70 |
0.75 |
0.86 |
| 5 |
E |
428 |
409 |
0.02 |
4.54 |
0.75 |
0.86 |
| 5 |
E |
428 |
409 |
0.02 |
4.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1166.0 cm
-1
Scaled (by 0.9554) Zero Point Vibrational Energy (zpe) 1114.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVTZ
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.437 |
0.316 |
| S2 |
0.000 |
-1.437 |
0.316 |
| S3 |
-1.437 |
0.000 |
-0.316 |
| S4 |
1.437 |
0.000 |
-0.316 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8735 | 2.1281 | 2.1281 |
S2 | 2.8735 | | 2.1281 | 2.1281 | S3 | 2.1281 | 2.1281 | | 2.8735 | S4 | 2.1281 | 2.1281 | 2.8735 | |
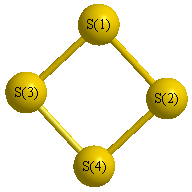 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.535 |
|
S1 |
S3 |
S4 |
47.535 |
| S2 |
S1 |
S3 |
47.535 |
|
S2 |
S4 |
S3 |
47.535 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULLultrafine/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.105 |
0.000 |
0.000 |
| y |
0.000 |
-49.067 |
0.000 |
| z |
0.000 |
0.000 |
-54.547 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.702 |
0.000 |
0.000 |
| y |
0.000 |
2.759 |
0.000 |
| z |
0.000 |
0.000 |
-5.461 |
|
| Polar |
|---|
| 3z2-r2 | -10.922 |
|---|
| x2-y2 | -0.038 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.447 |
0.000 |
0.000 |
| y |
0.000 |
11.443 |
0.000 |
| z |
0.000 |
0.000 |
6.618 |
<r2> (average value of r
2) Å
2
| <r2> |
170.310 |
| (<r2>)1/2 |
13.050 |
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULLultrafine/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.382936 |
| Energy at 298.15K | |
| HF Energy | -1592.060919 |
| Nuclear repulsion energy | 329.898774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
693 |
662 |
0.00 |
35.17 |
0.16 |
0.28 |
| 2 |
A1 |
249 |
238 |
0.00 |
3.27 |
0.73 |
0.84 |
| 3 |
A1 |
108 |
103 |
8.42 |
0.00 |
0.68 |
0.81 |
| 4 |
A2 |
232 |
222 |
0.00 |
0.00 |
0.75 |
0.86 |
| 5 |
B2 |
675 |
645 |
41.14 |
0.00 |
0.75 |
0.86 |
| 6 |
B2 |
318 |
304 |
0.00 |
12.42 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1137.4 cm
-1
Scaled (by 0.9554) Zero Point Vibrational Energy (zpe) 1086.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.298 |
0.957 |
| S2 |
0.000 |
-1.298 |
0.957 |
| S3 |
0.000 |
1.298 |
-0.957 |
| S4 |
0.000 |
-1.298 |
-0.957 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5963 | 1.9142 | 3.2256 |
S2 | 2.5963 | | 3.2256 | 1.9142 | S3 | 1.9142 | 3.2256 | | 2.5960 | S4 | 3.2256 | 1.9142 | 2.5960 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
89.996 |
|
S1 |
S3 |
S4 |
90.004 |
| S2 |
S1 |
S3 |
89.996 |
|
S2 |
S4 |
S3 |
90.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULLultrafine/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.000 |
|
|
|
| 2 |
S |
-0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.805 |
0.000 |
0.000 |
| y |
0.000 |
-52.956 |
0.000 |
| z |
0.000 |
0.000 |
-48.124 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.265 |
0.000 |
0.000 |
| y |
0.000 |
-2.992 |
0.000 |
| z |
0.000 |
0.000 |
4.256 |
|
| Polar |
|---|
| 3z2-r2 | 8.512 |
|---|
| x2-y2 | 1.151 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.773 |
0.000 |
0.000 |
| y |
0.000 |
15.651 |
0.000 |
| z |
0.000 |
0.000 |
12.672 |
<r2> (average value of r
2) Å
2
| <r2> |
198.298 |
| (<r2>)1/2 |
14.082 |
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULLultrafine/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.382936 |
| Energy at 298.15K | |
| HF Energy | -1592.060914 |
| Nuclear repulsion energy | 329.886335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
693 |
662 |
0.00 |
35.18 |
0.16 |
0.28 |
| 2 |
Ag |
249 |
238 |
0.00 |
3.30 |
0.73 |
0.84 |
| 3 |
Au |
232 |
222 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
675 |
645 |
41.11 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
108 |
103 |
8.41 |
0.00 |
0.00 |
0.00 |
| 6 |
B3g |
318 |
304 |
0.00 |
12.42 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1137.3 cm
-1
Scaled (by 0.9554) Zero Point Vibrational Energy (zpe) 1086.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVTZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.957 |
1.298 |
| S2 |
0.000 |
0.957 |
-1.298 |
| S3 |
0.000 |
-0.957 |
1.298 |
| S4 |
0.000 |
-0.957 |
-1.298 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5964 | 1.9142 | 3.2257 |
S2 | 2.5964 | | 3.2257 | 1.9142 | S3 | 1.9142 | 3.2257 | | 2.5964 | S4 | 3.2257 | 1.9142 | 2.5964 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
