Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -114.244721 |
| Energy at 298.15K | -114.246144 |
| HF Energy | -113.835598 |
| Nuclear repulsion energy | 30.858975 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3779 |
3588 |
104.91 |
|
|
|
| 2 |
A' |
2961 |
2811 |
127.91 |
|
|
|
| 3 |
A' |
1526 |
1449 |
15.12 |
|
|
|
| 4 |
A' |
1372 |
1302 |
70.15 |
|
|
|
| 5 |
A' |
1219 |
1157 |
140.12 |
|
|
|
| 6 |
A" |
1132 |
1075 |
117.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5994.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 5691.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.739 |
0.000 |
| O2 |
0.010 |
-0.569 |
0.000 |
| H3 |
-1.066 |
0.974 |
0.000 |
| H4 |
0.932 |
-0.854 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3080 | 1.1012 | 1.8408 |
O2 | 1.3080 | | 1.8811 | 0.9653 | H3 | 1.1012 | 1.8811 | | 2.7083 | H4 | 1.8408 | 0.9653 | 2.7083 | |
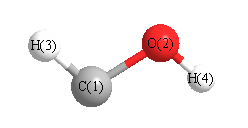 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
107.175 |
|
O2 |
C1 |
H3 |
102.330 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -114.236924 |
| Energy at 298.15K | -114.238340 |
| HF Energy | -113.827646 |
| Nuclear repulsion energy | 30.793636 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3675 |
3489 |
24.57 |
|
|
|
| 2 |
A' |
2875 |
2730 |
182.24 |
|
|
|
| 3 |
A' |
1501 |
1425 |
42.27 |
|
|
|
| 4 |
A' |
1378 |
1309 |
74.76 |
|
|
|
| 5 |
A' |
1244 |
1181 |
42.52 |
|
|
|
| 6 |
A" |
1062 |
1008 |
26.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5867.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 5570.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.122 |
0.739 |
0.000 |
| O2 |
0.122 |
-0.566 |
0.000 |
| H3 |
-0.939 |
1.059 |
0.000 |
| H4 |
-0.765 |
-0.960 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3050 | 1.1077 | 1.9164 |
O2 | 1.3050 | | 1.9408 | 0.9701 | H3 | 1.1077 | 1.9408 | | 2.0271 | H4 | 1.9164 | 0.9701 | 2.0271 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
113.969 |
|
O2 |
C1 |
H3 |
106.824 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
