Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -114.278631 |
| Energy at 298.15K | -114.280048 |
| HF Energy | -113.835182 |
| Nuclear repulsion energy | 30.697918 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3759 |
3620 |
|
|
|
|
| 2 |
A' |
2888 |
2781 |
|
|
|
|
| 3 |
A' |
1517 |
1461 |
|
|
|
|
| 4 |
A' |
1325 |
1276 |
|
|
|
|
| 5 |
A' |
1211 |
1166 |
|
|
|
|
| 6 |
A" |
1115 |
1073 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5907.6 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 5688.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.744 |
0.000 |
| O2 |
0.010 |
-0.573 |
0.000 |
| H3 |
-1.072 |
0.974 |
0.000 |
| H4 |
0.934 |
-0.855 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3168 | 1.1066 | 1.8464 |
O2 | 1.3168 | | 1.8883 | 0.9660 | H3 | 1.1066 | 1.8883 | | 2.7149 | H4 | 1.8464 | 0.9660 | 2.7149 | |
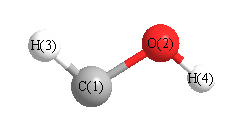 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
106.961 |
|
O2 |
C1 |
H3 |
102.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -114.271059 |
| Energy at 298.15K | -114.272468 |
| HF Energy | -113.827295 |
| Nuclear repulsion energy | 30.637182 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3658 |
3522 |
|
|
|
|
| 2 |
A' |
2799 |
2695 |
|
|
|
|
| 3 |
A' |
1488 |
1433 |
|
|
|
|
| 4 |
A' |
1337 |
1287 |
|
|
|
|
| 5 |
A' |
1230 |
1184 |
|
|
|
|
| 6 |
A" |
1043 |
1004 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5777.0 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 5562.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.122 |
0.743 |
0.000 |
| O2 |
0.122 |
-0.570 |
0.000 |
| H3 |
-0.945 |
1.060 |
0.000 |
| H4 |
-0.767 |
-0.959 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3136 | 1.1135 | 1.9209 |
O2 | 1.3136 | | 1.9486 | 0.9704 | H3 | 1.1135 | 1.9486 | | 2.0272 | H4 | 1.9209 | 0.9704 | 2.0272 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
113.647 |
|
O2 |
C1 |
H3 |
106.513 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
