Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.032353 |
| Energy at 298.15K | -117.036627 |
| HF Energy | -116.511496 |
| Nuclear repulsion energy | 64.913351 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3257 |
3174 |
|
|
|
|
| 2 |
A1 |
3164 |
3085 |
|
|
|
|
| 3 |
A1 |
3156 |
3076 |
|
|
|
|
| 4 |
A1 |
1533 |
1495 |
|
|
|
|
| 5 |
A1 |
1277 |
1245 |
|
|
|
|
| 6 |
A1 |
1039 |
1013 |
|
|
|
|
| 7 |
A1 |
428 |
417 |
|
|
|
|
| 8 |
A2 |
786 |
766 |
|
|
|
|
| 9 |
A2 |
560 |
546 |
|
|
|
|
| 10 |
B1 |
1011 |
986 |
|
|
|
|
| 11 |
B1 |
802 |
782 |
|
|
|
|
| 12 |
B1 |
534 |
520 |
|
|
|
|
| 13 |
B2 |
3253 |
3171 |
|
|
|
|
| 14 |
B2 |
3153 |
3074 |
|
|
|
|
| 15 |
B2 |
1518 |
1480 |
|
|
|
|
| 16 |
B2 |
1423 |
1387 |
|
|
|
|
| 17 |
B2 |
1185 |
1156 |
|
|
|
|
| 18 |
B2 |
939 |
916 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14509.3 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 14143.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.450 |
| H2 |
0.000 |
0.000 |
1.536 |
| C3 |
0.000 |
1.226 |
-0.199 |
| C4 |
0.000 |
-1.226 |
-0.199 |
| H5 |
0.000 |
2.156 |
0.353 |
| H6 |
0.000 |
-2.156 |
0.353 |
| H7 |
0.000 |
1.278 |
-1.281 |
| H8 |
0.000 |
-1.278 |
-1.281 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0857 | 1.3872 | 1.3872 | 2.1579 | 2.1579 | 2.1515 | 2.1515 |
H2 | 1.0857 | | 2.1241 | 2.1241 | 2.4589 | 2.4589 | 3.0929 | 3.0929 | C3 | 1.3872 | 2.1241 | | 2.4521 | 1.0811 | 3.4265 | 1.0833 | 2.7276 | C4 | 1.3872 | 2.1241 | 2.4521 | | 3.4265 | 1.0811 | 2.7276 | 1.0833 | H5 | 2.1579 | 2.4589 | 1.0811 | 3.4265 | | 4.3114 | 1.8548 | 3.8024 | H6 | 2.1579 | 2.4589 | 3.4265 | 1.0811 | 4.3114 | | 3.8024 | 1.8548 | H7 | 2.1515 | 3.0929 | 1.0833 | 2.7276 | 1.8548 | 3.8024 | | 2.5556 | H8 | 2.1515 | 3.0929 | 2.7276 | 1.0833 | 3.8024 | 1.8548 | 2.5556 | |
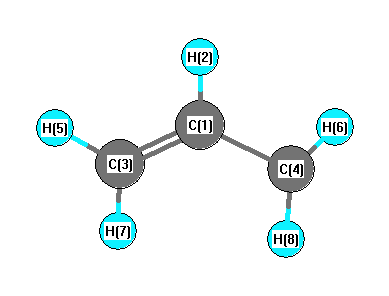 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.420 |
|
C1 |
C3 |
H7 |
120.629 |
| C1 |
C4 |
H6 |
121.420 |
|
C1 |
C4 |
H8 |
120.629 |
| H2 |
C1 |
C3 |
117.891 |
|
H2 |
C1 |
C4 |
117.891 |
| C3 |
C1 |
C4 |
124.219 |
|
H5 |
C3 |
H7 |
117.952 |
| H6 |
C4 |
H8 |
117.952 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.032353 |
| Energy at 298.15K | -117.036585 |
| HF Energy | -116.511486 |
| Nuclear repulsion energy | 64.910619 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3253 |
3171 |
|
|
|
|
| 2 |
A' |
3249 |
3167 |
|
|
|
|
| 3 |
A' |
3158 |
3078 |
|
|
|
|
| 4 |
A' |
3149 |
3070 |
|
|
|
|
| 5 |
A' |
3146 |
3067 |
|
|
|
|
| 6 |
A' |
1526 |
1487 |
|
|
|
|
| 7 |
A' |
1513 |
1475 |
|
|
|
|
| 8 |
A' |
1418 |
1383 |
|
|
|
|
| 9 |
A' |
1273 |
1241 |
|
|
|
|
| 10 |
A' |
1179 |
1149 |
|
|
|
|
| 11 |
A' |
1034 |
1008 |
|
|
|
|
| 12 |
A' |
926 |
903 |
|
|
|
|
| 13 |
A' |
415 |
404 |
|
|
|
|
| 14 |
A" |
1000 |
975 |
|
|
|
|
| 15 |
A" |
798 |
778 |
|
|
|
|
| 16 |
A" |
776 |
756 |
|
|
|
|
| 17 |
A" |
544 |
530 |
|
|
|
|
| 18 |
A" |
524 |
511 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14440.7 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 14076.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.451 |
0.000 |
| H2 |
-0.000 |
1.537 |
0.000 |
| C3 |
-1.226 |
-0.199 |
0.000 |
| C4 |
1.226 |
-0.199 |
0.000 |
| H5 |
-2.156 |
0.351 |
0.000 |
| H6 |
2.156 |
0.352 |
0.000 |
| H7 |
-1.277 |
-1.281 |
0.000 |
| H8 |
1.277 |
-1.281 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0859 | 1.3871 | 1.3871 | 2.1587 | 2.1583 | 2.1520 | 2.1519 |
H2 | 1.0859 | | 2.1246 | 2.1248 | 2.4606 | 2.4603 | 3.0938 | 3.0939 | C3 | 1.3871 | 2.1246 | | 2.4512 | 1.0812 | 3.4262 | 1.0836 | 2.7265 | C4 | 1.3871 | 2.1248 | 2.4512 | | 3.4264 | 1.0812 | 2.7269 | 1.0836 | H5 | 2.1587 | 2.4606 | 1.0812 | 3.4264 | | 4.3124 | 1.8541 | 3.8016 | H6 | 2.1583 | 2.4603 | 3.4262 | 1.0812 | 4.3124 | | 3.8019 | 1.8548 | H7 | 2.1520 | 3.0938 | 1.0836 | 2.7269 | 1.8541 | 3.8019 | | 2.5542 | H8 | 2.1519 | 3.0939 | 2.7265 | 1.0836 | 3.8016 | 1.8548 | 2.5542 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.490 |
|
C1 |
C3 |
H7 |
120.663 |
| C1 |
C4 |
H6 |
121.451 |
|
C1 |
C4 |
H8 |
120.637 |
| H2 |
C1 |
C3 |
117.916 |
|
H2 |
C1 |
C4 |
117.938 |
| C3 |
C1 |
C4 |
124.147 |
|
H5 |
C3 |
H7 |
117.847 |
| H6 |
C4 |
H8 |
117.912 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
