Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.014822 |
| Energy at 298.15K | -117.019226 |
| HF Energy | -116.511158 |
| Nuclear repulsion energy | 65.680638 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3320 |
3152 |
7.67 |
|
|
|
| 2 |
A1 |
3231 |
3068 |
0.08 |
|
|
|
| 3 |
A1 |
3219 |
3056 |
10.11 |
|
|
|
| 4 |
A1 |
1565 |
1486 |
3.64 |
|
|
|
| 5 |
A1 |
1317 |
1250 |
1.42 |
|
|
|
| 6 |
A1 |
1076 |
1022 |
0.02 |
|
|
|
| 7 |
A1 |
437 |
415 |
0.21 |
|
|
|
| 8 |
A2 |
863 |
820 |
0.00 |
|
|
|
| 9 |
A2 |
600 |
570 |
0.00 |
|
|
|
| 10 |
B1 |
1088 |
1033 |
21.99 |
|
|
|
| 11 |
B1 |
888 |
843 |
80.65 |
|
|
|
| 12 |
B1 |
575 |
546 |
15.96 |
|
|
|
| 13 |
B2 |
3315 |
3147 |
2.14 |
|
|
|
| 14 |
B2 |
3220 |
3057 |
6.05 |
|
|
|
| 15 |
B2 |
1530 |
1452 |
0.78 |
|
|
|
| 16 |
B2 |
1437 |
1365 |
8.99 |
|
|
|
| 17 |
B2 |
1145 |
1087 |
0.41 |
|
|
|
| 18 |
B2 |
955 |
907 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14891.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14137.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.443 |
| H2 |
0.000 |
0.000 |
1.521 |
| C3 |
0.000 |
1.210 |
-0.195 |
| C4 |
0.000 |
-1.210 |
-0.195 |
| H5 |
0.000 |
2.133 |
0.352 |
| H6 |
0.000 |
-2.133 |
0.352 |
| H7 |
0.000 |
1.261 |
-1.270 |
| H8 |
0.000 |
-1.261 |
-1.270 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0774 | 1.3679 | 1.3679 | 2.1350 | 2.1350 | 2.1269 | 2.1269 |
H2 | 1.0774 | | 2.0994 | 2.0994 | 2.4321 | 2.4321 | 3.0620 | 3.0620 | C3 | 1.3679 | 2.0994 | | 2.4194 | 1.0734 | 3.3873 | 1.0758 | 2.6939 | C4 | 1.3679 | 2.0994 | 2.4194 | | 3.3873 | 1.0734 | 2.6939 | 1.0758 | H5 | 2.1350 | 2.4321 | 1.0734 | 3.3873 | | 4.2661 | 1.8418 | 3.7614 | H6 | 2.1350 | 2.4321 | 3.3873 | 1.0734 | 4.2661 | | 3.7614 | 1.8418 | H7 | 2.1269 | 3.0620 | 1.0758 | 2.6939 | 1.8418 | 3.7614 | | 2.5212 | H8 | 2.1269 | 3.0620 | 2.6939 | 1.0758 | 3.7614 | 1.8418 | 2.5212 | |
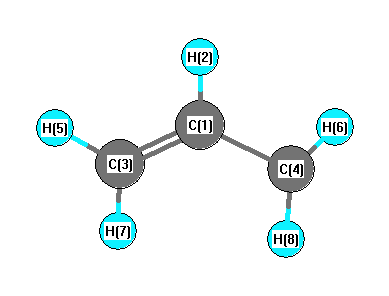 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.512 |
|
C1 |
C3 |
H7 |
120.534 |
| C1 |
C4 |
H6 |
121.512 |
|
C1 |
C4 |
H8 |
120.534 |
| H2 |
C1 |
C3 |
117.824 |
|
H2 |
C1 |
C4 |
117.824 |
| C3 |
C1 |
C4 |
124.351 |
|
H5 |
C3 |
H7 |
117.954 |
| H6 |
C4 |
H8 |
117.954 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.014822 |
| Energy at 298.15K | -117.019226 |
| HF Energy | -116.511148 |
| Nuclear repulsion energy | 65.685884 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3320 |
3152 |
7.69 |
|
|
|
| 2 |
A' |
3315 |
3147 |
2.13 |
|
|
|
| 3 |
A' |
3231 |
3068 |
0.04 |
|
|
|
| 4 |
A' |
3220 |
3057 |
9.78 |
|
|
|
| 5 |
A' |
3220 |
3057 |
6.40 |
|
|
|
| 6 |
A' |
1565 |
1486 |
3.66 |
|
|
|
| 7 |
A' |
1530 |
1452 |
0.78 |
|
|
|
| 8 |
A' |
1437 |
1365 |
9.00 |
|
|
|
| 9 |
A' |
1317 |
1250 |
1.43 |
|
|
|
| 10 |
A' |
1145 |
1087 |
0.40 |
|
|
|
| 11 |
A' |
1077 |
1022 |
0.02 |
|
|
|
| 12 |
A' |
955 |
907 |
0.08 |
|
|
|
| 13 |
A' |
438 |
415 |
0.21 |
|
|
|
| 14 |
A" |
1088 |
1033 |
21.98 |
|
|
|
| 15 |
A" |
888 |
843 |
80.63 |
|
|
|
| 16 |
A" |
863 |
820 |
0.04 |
|
|
|
| 17 |
A" |
600 |
570 |
0.00 |
|
|
|
| 18 |
A" |
575 |
546 |
15.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14891.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14138.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.443 |
0.000 |
| H2 |
0.000 |
1.521 |
0.000 |
| C3 |
-1.210 |
-0.196 |
0.000 |
| C4 |
1.209 |
-0.195 |
0.000 |
| H5 |
-2.133 |
0.351 |
0.000 |
| H6 |
2.133 |
0.352 |
0.000 |
| H7 |
-1.259 |
-1.270 |
0.000 |
| H8 |
1.260 |
-1.270 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0773 | 1.3681 | 1.3675 | 2.1353 | 2.1347 | 2.1265 | 2.1266 |
H2 | 1.0773 | | 2.0999 | 2.0990 | 2.4329 | 2.4316 | 3.0619 | 3.0616 | C3 | 1.3681 | 2.0999 | | 2.4190 | 1.0735 | 3.3870 | 1.0758 | 2.6932 | C4 | 1.3675 | 2.0990 | 2.4190 | | 3.3870 | 1.0734 | 2.6925 | 1.0758 | H5 | 2.1353 | 2.4329 | 1.0735 | 3.3870 | | 4.2660 | 1.8423 | 3.7607 | H6 | 2.1347 | 2.4316 | 3.3870 | 1.0734 | 4.2660 | | 3.7599 | 1.8418 | H7 | 2.1265 | 3.0619 | 1.0758 | 2.6925 | 1.8423 | 3.7599 | | 2.5192 | H8 | 2.1266 | 3.0616 | 2.6932 | 1.0758 | 3.7607 | 1.8418 | 2.5192 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.519 |
|
C1 |
C3 |
H7 |
120.482 |
| C1 |
C4 |
H6 |
121.515 |
|
C1 |
C4 |
H8 |
120.535 |
| H2 |
C1 |
C3 |
117.858 |
|
H2 |
C1 |
C4 |
117.819 |
| C3 |
C1 |
C4 |
124.324 |
|
H5 |
C3 |
H7 |
118.000 |
| H6 |
C4 |
H8 |
117.950 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
