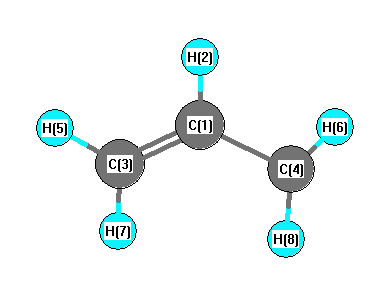
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3268 |
3128 |
15.64 |
|
|
|
| 2 |
A1 |
3170 |
3034 |
1.84 |
|
|
|
| 3 |
A1 |
3162 |
3027 |
14.44 |
|
|
|
| 4 |
A1 |
1521 |
1456 |
3.09 |
|
|
|
| 5 |
A1 |
1279 |
1225 |
1.29 |
|
|
|
| 6 |
A1 |
1045 |
1000 |
0.03 |
|
|
|
| 7 |
A1 |
416 |
398 |
0.12 |
|
|
|
| 8 |
A2 |
800 |
766 |
0.00 |
|
|
|
| 9 |
A2 |
558 |
534 |
0.00 |
|
|
|
| 10 |
B1 |
1022 |
978 |
19.27 |
|
|
|
| 11 |
B1 |
826 |
790 |
73.50 |
|
|
|
| 12 |
B1 |
533 |
510 |
15.51 |
|
|
|
| 13 |
B2 |
3266 |
3126 |
4.31 |
|
|
|
| 14 |
B2 |
3163 |
3028 |
7.36 |
|
|
|
| 15 |
B2 |
1519 |
1454 |
1.02 |
|
|
|
| 16 |
B2 |
1423 |
1362 |
6.16 |
|
|
|
| 17 |
B2 |
1214 |
1162 |
0.53 |
|
|
|
| 18 |
B2 |
932 |
892 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14557.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 13932.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.055 |
|
|
|
| 2 |
H |
0.142 |
|
|
|
| 3 |
C |
-0.303 |
|
|
|
| 4 |
C |
-0.303 |
|
|
|
| 5 |
H |
0.133 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.062 |
0.062 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.009 |
0.000 |
0.000 |
| y |
0.000 |
-17.845 |
0.000 |
| z |
0.000 |
0.000 |
-17.573 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.301 |
0.000 |
0.000 |
| y |
0.000 |
1.946 |
0.000 |
| z |
0.000 |
0.000 |
2.355 |
|
| Polar |
|---|
| 3z2-r2 | 4.709 |
|---|
| x2-y2 | -4.164 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.193 |
0.000 |
0.000 |
| y |
0.000 |
8.154 |
0.000 |
| z |
0.000 |
0.000 |
5.038 |
<r2> (average value of r
2) Å
2
| <r2> |
49.696 |
| (<r2>)1/2 |
7.050 |