return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (3-Penten-1-yne, (E)-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-193.692376
Energy at 298.15K-193.696816
HF Energy-192.816535
Nuclear repulsion energy138.761540
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3500 3323 69.38      
2 A' 3214 3052 13.00      
3 A' 3202 3040 4.81      
4 A' 3169 3009 9.81      
5 A' 3082 2926 18.78      
6 A' 2143 2035 0.32      
7 A' 1711 1624 1.41      
8 A' 1514 1437 11.41      
9 A' 1422 1350 1.74      
10 A' 1336 1268 0.98      
11 A' 1320 1253 3.23      
12 A' 1147 1089 0.03      
13 A' 1061 1008 9.57      
14 A' 917 870 6.76      
15 A' 654 621 41.80      
16 A' 537 510 2.05      
17 A' 383 364 2.09      
18 A' 165 156 1.29      
19 A" 3143 2984 10.07      
20 A" 1504 1428 7.09      
21 A" 1082 1027 1.02      
22 A" 1018 966 39.77      
23 A" 842 799 0.43      
24 A" 634 602 43.45      
25 A" 458 435 0.86      
26 A" 210 199 0.07      
27 A" 172 163 3.07      

Unscaled Zero Point Vibrational Energy (zpe) 19768.8 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18768.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.30914 0.07547 0.07230

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.321 -0.418 0.000
H2 -2.746 -0.904 0.875
H3 -2.746 -0.904 -0.875
H4 -2.627 0.623 0.000
C5 -0.839 -0.543 0.000
H6 -0.431 -1.542 0.000
C7 0.000 0.494 0.000
H8 -0.388 1.500 0.000
C9 1.414 0.361 0.000
C10 2.623 0.281 0.000
H11 3.675 0.180 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 C10 H11
C11.08691.08691.08491.48672.19892.49352.72303.81504.99316.0250
H21.08691.74921.76312.12852.55613.20293.47894.43505.56756.5699
H31.08691.74921.76312.12852.55613.20293.47894.43505.56756.5699
H41.08491.76311.76312.13403.08342.62972.40424.04895.26086.3167
C51.48672.12852.12852.13401.07901.33422.09282.42793.55924.5714
H62.19892.55612.55613.08341.07902.08083.04262.65033.55654.4517
C72.49353.20293.20292.62971.33422.08081.07881.42012.63183.6880
H82.72303.47893.47892.40422.09283.04261.07882.13203.24904.2720
C93.81504.43504.43504.04892.42792.65031.42012.13201.21202.2680
C104.99315.56755.56755.26083.55923.55652.63183.24901.21201.0563
H116.02506.56996.56996.31674.57144.45173.68804.27202.26801.0563

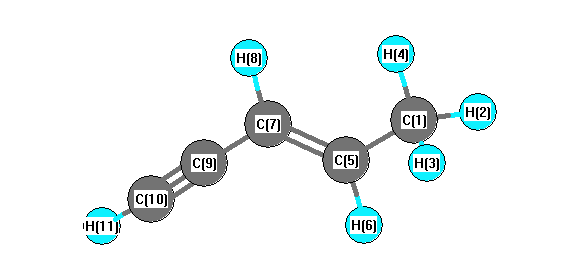
picture of 3-Penten-1-yne, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 117.082 C1 C5 C7 124.151
H2 C1 H3 107.154 H2 C1 H4 108.546
H2 C1 C5 110.634 H3 C1 H4 108.546
H3 C1 C5 110.634 H4 C1 C5 111.203
C5 C7 H8 119.924 C5 C7 C9 123.619
H6 C5 C7 118.767 C7 C9 C10 178.435
H8 C7 C9 116.457 C9 C10 H11 178.308
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability