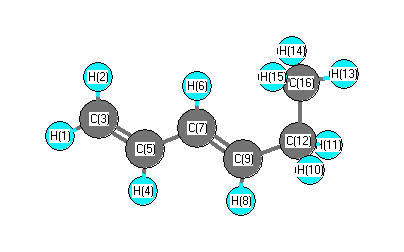
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3252 |
3112 |
13.13 |
|
|
|
| 2 |
A' |
3176 |
3039 |
20.23 |
|
|
|
| 3 |
A' |
3163 |
3028 |
2.89 |
|
|
|
| 4 |
A' |
3154 |
3019 |
12.29 |
|
|
|
| 5 |
A' |
3145 |
3010 |
16.81 |
|
|
|
| 6 |
A' |
3135 |
3001 |
32.47 |
|
|
|
| 7 |
A' |
3058 |
2927 |
20.01 |
|
|
|
| 8 |
A' |
3027 |
2897 |
29.11 |
|
|
|
| 9 |
A' |
1744 |
1670 |
24.00 |
|
|
|
| 10 |
A' |
1689 |
1616 |
6.52 |
|
|
|
| 11 |
A' |
1511 |
1447 |
5.41 |
|
|
|
| 12 |
A' |
1470 |
1407 |
6.53 |
|
|
|
| 13 |
A' |
1455 |
1392 |
4.26 |
|
|
|
| 14 |
A' |
1415 |
1355 |
6.15 |
|
|
|
| 15 |
A' |
1380 |
1321 |
10.76 |
|
|
|
| 16 |
A' |
1349 |
1291 |
3.66 |
|
|
|
| 17 |
A' |
1328 |
1271 |
2.02 |
|
|
|
| 18 |
A' |
1306 |
1250 |
0.08 |
|
|
|
| 19 |
A' |
1208 |
1156 |
4.23 |
|
|
|
| 20 |
A' |
1110 |
1062 |
4.12 |
|
|
|
| 21 |
A' |
1043 |
999 |
4.40 |
|
|
|
| 22 |
A' |
958 |
917 |
1.08 |
|
|
|
| 23 |
A' |
905 |
866 |
3.26 |
|
|
|
| 24 |
A' |
570 |
546 |
3.05 |
|
|
|
| 25 |
A' |
470 |
450 |
0.37 |
|
|
|
| 26 |
A' |
321 |
307 |
0.42 |
|
|
|
| 27 |
A' |
159 |
152 |
0.99 |
|
|
|
| 28 |
A" |
3128 |
2993 |
28.35 |
|
|
|
| 29 |
A" |
3046 |
2916 |
13.47 |
|
|
|
| 30 |
A" |
1504 |
1439 |
7.84 |
|
|
|
| 31 |
A" |
1287 |
1232 |
0.20 |
|
|
|
| 32 |
A" |
1102 |
1054 |
1.43 |
|
|
|
| 33 |
A" |
1055 |
1010 |
39.45 |
|
|
|
| 34 |
A" |
995 |
953 |
10.54 |
|
|
|
| 35 |
A" |
939 |
898 |
35.84 |
|
|
|
| 36 |
A" |
886 |
848 |
1.96 |
|
|
|
| 37 |
A" |
765 |
733 |
7.27 |
|
|
|
| 38 |
A" |
643 |
616 |
1.17 |
|
|
|
| 39 |
A" |
324 |
311 |
1.25 |
|
|
|
| 40 |
A" |
259 |
247 |
0.72 |
|
|
|
| 41 |
A" |
167 |
160 |
1.11 |
|
|
|
| 42 |
A" |
122 |
117 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31362.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30016.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.130 |
|
|
|
| 2 |
H |
0.123 |
|
|
|
| 3 |
C |
-0.343 |
|
|
|
| 4 |
H |
0.125 |
|
|
|
| 5 |
C |
-0.062 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
C |
-0.118 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
C |
-0.153 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
C |
-0.165 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
| 15 |
H |
0.123 |
|
|
|
| 16 |
C |
-0.389 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.159 |
-0.768 |
0.000 |
0.784 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.281 |
0.296 |
0.000 |
| y |
0.296 |
-36.413 |
0.000 |
| z |
0.000 |
0.000 |
-41.174 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.512 |
0.296 |
0.000 |
| y |
0.296 |
2.315 |
0.000 |
| z |
0.000 |
0.000 |
-4.827 |
|
| Polar |
|---|
| 3z2-r2 | -9.654 |
|---|
| x2-y2 | 0.132 |
|---|
| xy | 0.296 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.351 |
1.667 |
0.000 |
| y |
1.667 |
16.979 |
0.000 |
| z |
0.000 |
0.000 |
6.956 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |