Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.716145 |
| Energy at 298.15K | -133.721653 |
| HF Energy | -133.126071 |
| Nuclear repulsion energy | 71.245551 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3500 |
3323 |
0.47 |
|
|
|
| 2 |
A' |
3199 |
3037 |
6.72 |
|
|
|
| 3 |
A' |
3095 |
2939 |
23.57 |
|
|
|
| 4 |
A' |
3087 |
2931 |
35.70 |
|
|
|
| 5 |
A' |
1718 |
1631 |
45.10 |
|
|
|
| 6 |
A' |
1497 |
1422 |
12.07 |
|
|
|
| 7 |
A' |
1443 |
1370 |
19.92 |
|
|
|
| 8 |
A' |
1403 |
1332 |
10.90 |
|
|
|
| 9 |
A' |
1287 |
1222 |
33.00 |
|
|
|
| 10 |
A' |
1074 |
1019 |
32.98 |
|
|
|
| 11 |
A' |
947 |
899 |
5.30 |
|
|
|
| 12 |
A' |
490 |
465 |
17.43 |
|
|
|
| 13 |
A" |
3156 |
2996 |
8.93 |
|
|
|
| 14 |
A" |
1506 |
1430 |
8.46 |
|
|
|
| 15 |
A" |
1148 |
1089 |
3.63 |
|
|
|
| 16 |
A" |
1092 |
1036 |
13.06 |
|
|
|
| 17 |
A" |
698 |
662 |
51.41 |
|
|
|
| 18 |
A" |
202 |
192 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15270.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14498.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.032 |
-0.623 |
0.000 |
| C2 |
0.000 |
0.448 |
0.000 |
| N3 |
1.239 |
0.174 |
0.000 |
| H4 |
-0.551 |
-1.594 |
0.000 |
| H5 |
-1.668 |
-0.531 |
0.875 |
| H6 |
-1.668 |
-0.531 |
-0.875 |
| H7 |
-0.384 |
1.467 |
0.000 |
| H8 |
1.791 |
1.026 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4870 | 2.4061 | 1.0828 | 1.0864 | 1.0864 | 2.1887 | 3.2689 |
C2 | 1.4870 | | 1.2685 | 2.1146 | 2.1231 | 2.1231 | 1.0895 | 1.8817 | N3 | 2.4061 | 1.2685 | | 2.5156 | 3.1167 | 3.1167 | 2.0749 | 1.0155 | H4 | 1.0828 | 2.1146 | 2.5156 | | 1.7732 | 1.7732 | 3.0657 | 3.5141 | H5 | 1.0864 | 2.1231 | 3.1167 | 1.7732 | | 1.7507 | 2.5316 | 3.8930 | H6 | 1.0864 | 2.1231 | 3.1167 | 1.7732 | 1.7507 | | 2.5316 | 3.8930 | H7 | 2.1887 | 1.0895 | 2.0749 | 3.0657 | 2.5316 | 2.5316 | | 2.2187 | H8 | 3.2689 | 1.8817 | 1.0155 | 3.5141 | 3.8930 | 3.8930 | 2.2187 | |
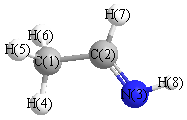 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.459 |
|
C1 |
C2 |
H7 |
115.446 |
| C2 |
C1 |
H4 |
109.742 |
|
C2 |
C1 |
H5 |
110.202 |
| C2 |
C1 |
H6 |
110.202 |
|
C2 |
N3 |
H8 |
110.463 |
| N3 |
C2 |
H7 |
123.095 |
|
H4 |
C1 |
H5 |
109.653 |
| H4 |
C1 |
H6 |
109.653 |
|
H5 |
C1 |
H6 |
107.354 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.715018 |
| Energy at 298.15K | -133.720515 |
| HF Energy | -133.124842 |
| Nuclear repulsion energy | 71.217067 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3463 |
3288 |
2.44 |
|
|
|
| 2 |
A' |
3178 |
3017 |
21.41 |
|
|
|
| 3 |
A' |
3144 |
2985 |
18.27 |
|
|
|
| 4 |
A' |
3088 |
2931 |
6.24 |
|
|
|
| 5 |
A' |
1713 |
1627 |
42.76 |
|
|
|
| 6 |
A' |
1504 |
1428 |
25.56 |
|
|
|
| 7 |
A' |
1444 |
1371 |
18.93 |
|
|
|
| 8 |
A' |
1410 |
1338 |
7.35 |
|
|
|
| 9 |
A' |
1287 |
1222 |
55.44 |
|
|
|
| 10 |
A' |
1081 |
1026 |
23.01 |
|
|
|
| 11 |
A' |
930 |
883 |
3.89 |
|
|
|
| 12 |
A' |
493 |
468 |
8.11 |
|
|
|
| 13 |
A" |
3157 |
2997 |
6.67 |
|
|
|
| 14 |
A" |
1501 |
1425 |
9.10 |
|
|
|
| 15 |
A" |
1171 |
1112 |
42.85 |
|
|
|
| 16 |
A" |
1085 |
1031 |
8.92 |
|
|
|
| 17 |
A" |
706 |
670 |
8.49 |
|
|
|
| 18 |
A" |
191 |
181 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15272.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14499.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.019 |
-0.607 |
0.000 |
| C2 |
0.000 |
0.481 |
0.000 |
| N3 |
1.266 |
0.360 |
0.000 |
| H4 |
-0.548 |
-1.585 |
0.000 |
| H5 |
-1.659 |
-0.523 |
0.874 |
| H6 |
-1.659 |
-0.523 |
-0.874 |
| H7 |
-0.390 |
1.493 |
0.000 |
| H8 |
1.509 |
-0.630 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4908 | 2.4810 | 1.0849 | 1.0867 | 1.0867 | 2.1924 | 2.5279 |
C2 | 1.4908 | | 1.2716 | 2.1374 | 2.1271 | 2.1271 | 1.0843 | 1.8741 | N3 | 2.4810 | 1.2716 | | 2.6596 | 3.1779 | 3.1779 | 2.0060 | 1.0198 | H4 | 1.0849 | 2.1374 | 2.6596 | | 1.7678 | 1.7678 | 3.0818 | 2.2680 | H5 | 1.0867 | 2.1271 | 3.1779 | 1.7678 | | 1.7479 | 2.5376 | 3.2882 | H6 | 1.0867 | 2.1271 | 3.1779 | 1.7678 | 1.7479 | | 2.5376 | 3.2882 | H7 | 2.1924 | 1.0843 | 2.0060 | 3.0818 | 2.5376 | 2.5376 | | 2.8484 | H8 | 2.5279 | 1.8741 | 1.0198 | 2.2680 | 3.2882 | 3.2882 | 2.8484 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.650 |
|
C1 |
C2 |
H7 |
115.828 |
| C2 |
C1 |
H4 |
111.188 |
|
C2 |
C1 |
H5 |
110.247 |
| C2 |
C1 |
H6 |
110.247 |
|
C2 |
N3 |
H8 |
109.255 |
| N3 |
C2 |
H7 |
116.522 |
|
H4 |
C1 |
H5 |
108.990 |
| H4 |
C1 |
H6 |
108.990 |
|
H5 |
C1 |
H6 |
107.074 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
