Jump to
S1C2
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.768956 |
| Energy at 298.15K | -133.774434 |
| HF Energy | -133.124487 |
| Nuclear repulsion energy | 70.992771 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3419 |
3292 |
|
|
|
|
| 2 |
A' |
3132 |
3016 |
|
|
|
|
| 3 |
A' |
3107 |
2991 |
|
|
|
|
| 4 |
A' |
3052 |
2939 |
|
|
|
|
| 5 |
A' |
1693 |
1630 |
|
|
|
|
| 6 |
A' |
1497 |
1441 |
|
|
|
|
| 7 |
A' |
1438 |
1385 |
|
|
|
|
| 8 |
A' |
1405 |
1353 |
|
|
|
|
| 9 |
A' |
1286 |
1238 |
|
|
|
|
| 10 |
A' |
1074 |
1034 |
|
|
|
|
| 11 |
A' |
920 |
885 |
|
|
|
|
| 12 |
A' |
488 |
470 |
|
|
|
|
| 13 |
A" |
3111 |
2996 |
|
|
|
|
| 14 |
A" |
1492 |
1437 |
|
|
|
|
| 15 |
A" |
1160 |
1117 |
|
|
|
|
| 16 |
A" |
1079 |
1039 |
|
|
|
|
| 17 |
A" |
700 |
674 |
|
|
|
|
| 18 |
A" |
187 |
180 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15119.4 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 14558.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.022 |
-0.610 |
0.000 |
| C2 |
0.000 |
0.484 |
0.000 |
| N3 |
1.270 |
0.362 |
0.000 |
| H4 |
-0.548 |
-1.589 |
0.000 |
| H5 |
-1.664 |
-0.527 |
0.877 |
| H6 |
-1.664 |
-0.527 |
-0.877 |
| H7 |
-0.393 |
1.497 |
0.000 |
| H8 |
1.512 |
-0.631 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4969 | 2.4895 | 1.0879 | 1.0895 | 1.0895 | 2.1984 | 2.5340 |
C2 | 1.4969 | | 1.2756 | 2.1442 | 2.1351 | 2.1351 | 1.0863 | 1.8783 | N3 | 2.4895 | 1.2756 | | 2.6669 | 3.1882 | 3.1882 | 2.0128 | 1.0222 | H4 | 1.0879 | 2.1442 | 2.6669 | | 1.7722 | 1.7722 | 3.0896 | 2.2719 | H5 | 1.0895 | 2.1351 | 3.1882 | 1.7722 | | 1.7533 | 2.5454 | 3.2959 | H6 | 1.0895 | 2.1351 | 3.1882 | 1.7722 | 1.7533 | | 2.5454 | 3.2959 | H7 | 2.1984 | 1.0863 | 2.0128 | 3.0896 | 2.5454 | 2.5454 | | 2.8556 | H8 | 2.5340 | 1.8783 | 1.0222 | 2.2719 | 3.2959 | 3.2959 | 2.8556 | |
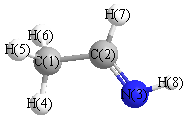 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.588 |
|
C1 |
C2 |
H7 |
115.732 |
| C2 |
C1 |
H4 |
111.114 |
|
C2 |
C1 |
H5 |
110.284 |
| C2 |
C1 |
H6 |
110.284 |
|
C2 |
N3 |
H8 |
109.171 |
| N3 |
C2 |
H7 |
116.680 |
|
H4 |
C1 |
H5 |
108.956 |
| H4 |
C1 |
H6 |
108.956 |
|
H5 |
C1 |
H6 |
107.148 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
