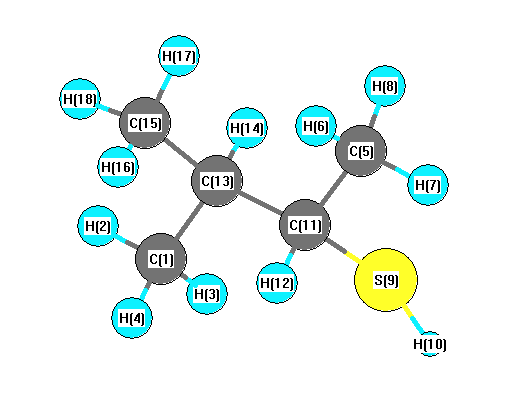
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3143 |
3009 |
24.37 |
|
|
|
| 2 |
A |
3137 |
3002 |
30.01 |
|
|
|
| 3 |
A |
3134 |
3000 |
21.51 |
|
|
|
| 4 |
A |
3128 |
2994 |
28.98 |
|
|
|
| 5 |
A |
3123 |
2989 |
50.78 |
|
|
|
| 6 |
A |
3117 |
2984 |
1.50 |
|
|
|
| 7 |
A |
3062 |
2931 |
29.86 |
|
|
|
| 8 |
A |
3057 |
2926 |
17.56 |
|
|
|
| 9 |
A |
3051 |
2920 |
22.21 |
|
|
|
| 10 |
A |
3048 |
2918 |
15.40 |
|
|
|
| 11 |
A |
3034 |
2904 |
2.34 |
|
|
|
| 12 |
A |
2717 |
2600 |
5.70 |
|
|
|
| 13 |
A |
1521 |
1456 |
0.96 |
|
|
|
| 14 |
A |
1503 |
1439 |
8.74 |
|
|
|
| 15 |
A |
1498 |
1433 |
20.55 |
|
|
|
| 16 |
A |
1488 |
1424 |
2.21 |
|
|
|
| 17 |
A |
1484 |
1420 |
2.93 |
|
|
|
| 18 |
A |
1482 |
1418 |
1.35 |
|
|
|
| 19 |
A |
1416 |
1355 |
6.47 |
|
|
|
| 20 |
A |
1409 |
1349 |
10.63 |
|
|
|
| 21 |
A |
1397 |
1337 |
5.06 |
|
|
|
| 22 |
A |
1379 |
1319 |
1.19 |
|
|
|
| 23 |
A |
1346 |
1288 |
0.39 |
|
|
|
| 24 |
A |
1318 |
1261 |
2.78 |
|
|
|
| 25 |
A |
1246 |
1192 |
21.63 |
|
|
|
| 26 |
A |
1203 |
1152 |
2.56 |
|
|
|
| 27 |
A |
1179 |
1128 |
6.93 |
|
|
|
| 28 |
A |
1160 |
1111 |
0.82 |
|
|
|
| 29 |
A |
1094 |
1047 |
1.12 |
|
|
|
| 30 |
A |
1047 |
1002 |
9.59 |
|
|
|
| 31 |
A |
983 |
941 |
2.79 |
|
|
|
| 32 |
A |
970 |
928 |
0.76 |
|
|
|
| 33 |
A |
934 |
894 |
1.80 |
|
|
|
| 34 |
A |
923 |
884 |
1.15 |
|
|
|
| 35 |
A |
895 |
857 |
2.62 |
|
|
|
| 36 |
A |
800 |
766 |
2.83 |
|
|
|
| 37 |
A |
700 |
670 |
1.76 |
|
|
|
| 38 |
A |
469 |
449 |
0.35 |
|
|
|
| 39 |
A |
395 |
378 |
0.38 |
|
|
|
| 40 |
A |
375 |
359 |
0.37 |
|
|
|
| 41 |
A |
365 |
349 |
0.12 |
|
|
|
| 42 |
A |
323 |
309 |
0.94 |
|
|
|
| 43 |
A |
247 |
236 |
10.87 |
|
|
|
| 44 |
A |
242 |
232 |
1.57 |
|
|
|
| 45 |
A |
217 |
207 |
0.13 |
|
|
|
| 46 |
A |
205 |
196 |
2.99 |
|
|
|
| 47 |
A |
187 |
179 |
0.99 |
|
|
|
| 48 |
A |
62 |
59 |
0.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35104.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 33598.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.360 |
|
|
|
| 2 |
H |
0.114 |
|
|
|
| 3 |
H |
0.134 |
|
|
|
| 4 |
H |
0.116 |
|
|
|
| 5 |
C |
-0.378 |
|
|
|
| 6 |
H |
0.121 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
S |
-0.198 |
|
|
|
| 10 |
H |
0.105 |
|
|
|
| 11 |
C |
-0.088 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
C |
-0.097 |
|
|
|
| 14 |
H |
0.144 |
|
|
|
| 15 |
C |
-0.385 |
|
|
|
| 16 |
H |
0.120 |
|
|
|
| 17 |
H |
0.126 |
|
|
|
| 18 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.920 |
1.266 |
0.494 |
1.641 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.960 |
2.565 |
1.458 |
| y |
2.565 |
-48.046 |
0.748 |
| z |
1.458 |
0.748 |
-48.027 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.076 |
2.565 |
1.458 |
| y |
2.565 |
-0.552 |
0.748 |
| z |
1.458 |
0.748 |
-0.524 |
|
| Polar |
|---|
| 3z2-r2 | -1.049 |
|---|
| x2-y2 | 1.085 |
|---|
| xy | 2.565 |
|---|
| xz | 1.458 |
|---|
| yz | 0.748 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.525 |
-0.244 |
0.010 |
| y |
-0.244 |
12.167 |
0.179 |
| z |
0.010 |
0.179 |
9.629 |
<r2> (average value of r
2) Å
2
| <r2> |
246.168 |
| (<r2>)1/2 |
15.690 |