Jump to
S2C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -498.836085 |
| Energy at 298.15K | -498.835956 |
| HF Energy | -498.836085 |
| Nuclear repulsion energy | 38.883021 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.045 |
1.188 |
0.000 |
| Cl2 |
0.045 |
-0.502 |
0.000 |
| H3 |
-1.040 |
1.415 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.6902 | 1.1083 |
Cl2 | 1.6902 | | 2.2031 | H3 | 1.1083 | 2.2031 | |
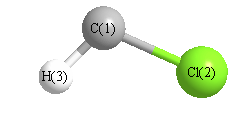 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
101.839 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.179 |
|
|
|
| 2 |
Cl |
0.050 |
|
|
|
| 3 |
H |
0.130 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.426 |
-0.669 |
0.000 |
1.575 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.382 |
-2.581 |
0.000 |
| y |
-2.581 |
-19.863 |
0.000 |
| z |
0.000 |
0.000 |
-17.936 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.482 |
-2.581 |
0.000 |
| y |
-2.581 |
-1.204 |
0.000 |
| z |
0.000 |
0.000 |
1.687 |
|
| Polar |
|---|
| 3z2-r2 | 3.373 |
|---|
| x2-y2 | 0.481 |
|---|
| xy | -2.581 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.924 |
-0.244 |
0.000 |
| y |
-0.244 |
5.043 |
0.000 |
| z |
0.000 |
0.000 |
2.468 |
<r2> (average value of r
2) Å
2
| <r2> |
27.791 |
| (<r2>)1/2 |
5.272 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -498.831224 |
| Energy at 298.15K | -498.831076 |
| HF Energy | -498.831224 |
| Nuclear repulsion energy | 39.273265 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.036 |
1.143 |
0.000 |
| Cl2 |
0.036 |
-0.509 |
0.000 |
| H3 |
-0.831 |
1.789 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.6520 | 1.0810 |
Cl2 | 1.6520 | | 2.4557 | H3 | 1.0810 | 2.4557 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
126.662 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.201 |
|
|
|
| 2 |
Cl |
0.030 |
|
|
|
| 3 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.736 |
0.250 |
0.000 |
0.777 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.313 |
-1.468 |
0.000 |
| y |
-1.468 |
-17.038 |
0.000 |
| z |
0.000 |
0.000 |
-19.442 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.073 |
-1.468 |
0.000 |
| y |
-1.468 |
1.840 |
0.000 |
| z |
0.000 |
0.000 |
-1.767 |
|
| Polar |
|---|
| 3z2-r2 | -3.534 |
|---|
| x2-y2 | -1.275 |
|---|
| xy | -1.468 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.539 |
-0.164 |
0.000 |
| y |
-0.164 |
4.808 |
0.000 |
| z |
0.000 |
0.000 |
2.288 |
<r2> (average value of r
2) Å
2
| <r2> |
27.570 |
| (<r2>)1/2 |
5.251 |
