Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -193.430494 |
| Energy at 298.15K | |
| HF Energy | -193.430494 |
| Nuclear repulsion energy | 149.745140 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3272 |
3132 |
0.18 |
|
|
|
| 2 |
A1 |
3264 |
3124 |
7.91 |
|
|
|
| 3 |
A1 |
3228 |
3089 |
1.21 |
|
|
|
| 4 |
A1 |
1598 |
1529 |
0.01 |
|
|
|
| 5 |
A1 |
1474 |
1410 |
2.35 |
|
|
|
| 6 |
A1 |
1157 |
1107 |
0.04 |
|
|
|
| 7 |
A1 |
1091 |
1045 |
8.34 |
|
|
|
| 8 |
A1 |
1050 |
1005 |
12.13 |
|
|
|
| 9 |
A1 |
842 |
806 |
0.28 |
|
|
|
| 10 |
A2 |
921 |
881 |
0.00 |
|
|
|
| 11 |
A2 |
833 |
797 |
0.00 |
|
|
|
| 12 |
A2 |
526 |
503 |
0.00 |
|
|
|
| 13 |
B1 |
901 |
862 |
6.40 |
|
|
|
| 14 |
B1 |
732 |
700 |
1.53 |
|
|
|
| 15 |
B1 |
687 |
658 |
100.17 |
|
|
|
| 16 |
B1 |
486 |
466 |
10.26 |
|
|
|
| 17 |
B2 |
3246 |
3107 |
13.61 |
|
|
|
| 18 |
B2 |
3229 |
3091 |
8.58 |
|
|
|
| 19 |
B2 |
1413 |
1352 |
40.56 |
|
|
|
| 20 |
B2 |
1299 |
1243 |
0.01 |
|
|
|
| 21 |
B2 |
1222 |
1169 |
2.21 |
|
|
|
| 22 |
B2 |
983 |
941 |
0.02 |
|
|
|
| 23 |
B2 |
925 |
885 |
4.36 |
|
|
|
| 24 |
B2 |
54i |
52i |
8.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17160.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 16424.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.216 |
| C2 |
0.000 |
1.115 |
0.390 |
| C3 |
0.000 |
0.675 |
-0.999 |
| C4 |
0.000 |
-0.675 |
-0.999 |
| C5 |
0.000 |
-1.115 |
0.390 |
| H6 |
0.000 |
0.000 |
2.292 |
| H7 |
0.000 |
2.145 |
0.712 |
| H8 |
0.000 |
1.327 |
-1.855 |
| H9 |
0.000 |
-1.327 |
-1.855 |
| H10 |
0.000 |
-2.145 |
0.712 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3878 | 2.3157 | 2.3157 | 1.3878 | 1.0753 | 2.2032 | 3.3459 | 3.3459 | 2.2032 |
C2 | 1.3878 | | 1.4573 | 2.2657 | 2.2304 | 2.2043 | 1.0785 | 2.2555 | 3.3175 | 3.2757 | C3 | 2.3157 | 1.4573 | | 1.3493 | 2.2657 | 3.3590 | 2.2556 | 1.0764 | 2.1770 | 3.2978 | C4 | 2.3157 | 2.2657 | 1.3493 | | 1.4573 | 3.3590 | 3.2978 | 2.1770 | 1.0764 | 2.2556 | C5 | 1.3878 | 2.2304 | 2.2657 | 1.4573 | | 2.2043 | 3.2757 | 3.3175 | 2.2555 | 1.0785 | H6 | 1.0753 | 2.2043 | 3.3590 | 3.3590 | 2.2043 | | 2.6637 | 4.3540 | 4.3540 | 2.6637 | H7 | 2.2032 | 1.0785 | 2.2556 | 3.2978 | 3.2757 | 2.6637 | | 2.6942 | 4.3176 | 4.2894 | H8 | 3.3459 | 2.2555 | 1.0764 | 2.1770 | 3.3175 | 4.3540 | 2.6942 | | 2.6537 | 4.3176 | H9 | 3.3459 | 3.3175 | 2.1770 | 1.0764 | 2.2555 | 4.3540 | 4.3176 | 2.6537 | | 2.6942 | H10 | 2.2032 | 3.2757 | 3.2978 | 2.2556 | 1.0785 | 2.6637 | 4.2894 | 4.3176 | 2.6942 | |
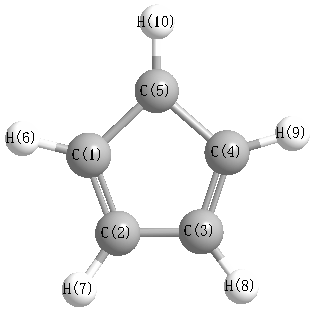 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.932 |
|
C1 |
C2 |
H7 |
126.124 |
| C1 |
C5 |
C4 |
108.932 |
|
C1 |
C5 |
H10 |
126.124 |
| C2 |
C1 |
C5 |
106.944 |
|
C2 |
C1 |
H6 |
126.528 |
| C2 |
C3 |
C4 |
107.596 |
|
C2 |
C3 |
H8 |
125.113 |
| C3 |
C2 |
H7 |
124.943 |
|
C3 |
C4 |
C5 |
107.596 |
| C3 |
C4 |
H9 |
127.292 |
|
C4 |
C3 |
H8 |
127.292 |
| C4 |
C5 |
H10 |
124.943 |
|
C5 |
C1 |
H6 |
126.528 |
| C5 |
C4 |
H9 |
125.113 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.213 |
|
|
|
| 2 |
C |
-0.078 |
|
|
|
| 3 |
C |
-0.165 |
|
|
|
| 4 |
C |
-0.165 |
|
|
|
| 5 |
C |
-0.078 |
|
|
|
| 6 |
H |
0.139 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.061 |
0.061 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.907 |
0.000 |
0.000 |
| y |
0.000 |
-25.491 |
0.000 |
| z |
0.000 |
0.000 |
-27.494 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.414 |
0.000 |
0.000 |
| y |
0.000 |
4.709 |
0.000 |
| z |
0.000 |
0.000 |
1.705 |
|
| Polar |
|---|
| 3z2-r2 | 3.409 |
|---|
| x2-y2 | -7.415 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.384 |
0.000 |
0.000 |
| y |
0.000 |
9.370 |
0.000 |
| z |
0.000 |
0.000 |
8.631 |
<r2> (average value of r
2) Å
2
| <r2> |
86.816 |
| (<r2>)1/2 |
9.317 |
