return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-118.227265
Energy at 298.15K 
HF Energy-117.675760
Nuclear repulsion energy76.473205
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3320 3152 8.48      
2 A 3229 3065 12.26      
3 A 3171 3011 22.29      
4 A 3165 3005 26.34      
5 A 3095 2938 27.80      
6 A 3093 2936 9.10      
7 A 3033 2879 23.28      
8 A 1535 1457 3.31      
9 A 1528 1450 6.71      
10 A 1506 1430 1.15      
11 A 1498 1422 2.97      
12 A 1425 1352 3.10      
13 A 1382 1312 1.44      
14 A 1293 1228 0.11      
15 A 1197 1136 0.21      
16 A 1123 1066 0.18      
17 A 1070 1016 1.03      
18 A 944 896 1.40      
19 A 911 865 0.18      
20 A 771 732 0.62      
21 A 495 470 47.19      
22 A 376 357 1.50      
23 A 273 259 0.02      
24 A 120 114 0.98      

Unscaled Zero Point Vibrational Energy (zpe) 19775.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18774.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.10738 0.30618 0.26415

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.287 -0.295 -0.030
C2 0.078 0.557 0.041
C3 -1.211 -0.242 -0.030
H4 -2.082 0.403 0.024
H5 -1.262 -0.804 -0.957
H6 -1.266 -0.951 0.791
H7 0.087 1.137 0.968
H8 0.102 1.294 -0.759
H9 2.254 0.133 -0.219
H10 1.241 -1.329 0.262

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.48122.49923.44112.76052.76132.11832.11321.07341.0744
C21.48121.51862.16522.15512.15501.09321.08842.23152.2262
C32.49921.51861.08491.08531.08602.14072.14863.49002.6977
H43.44112.16521.08491.75781.75712.47632.48464.35023.7542
H52.76052.15511.08531.75781.75473.04832.51013.71262.8333
H62.76132.15501.08601.75711.75472.49453.05223.81872.5901
H72.11831.09322.14072.47633.04832.49451.73402.66632.8122
H82.11321.08842.14862.48462.51013.05221.73402.50443.0363
H91.07342.23153.49004.35023.71263.81872.66632.50441.8415
H101.07442.22622.69773.75422.83332.59012.81223.03631.8415

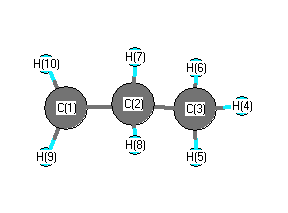
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.838 C1 C2 H7 109.821
C1 C2 H8 109.700 C2 C1 H9 120.912
C2 C1 H10 120.332 C2 C3 H4 111.454
C2 C3 H5 110.613 C2 C3 H6 110.564
C3 C2 H7 109.011 C3 C2 H8 109.911
H4 C3 H5 108.186 H4 C3 H6 108.076
H5 C3 H6 107.823 H7 C2 H8 105.282
H9 C1 H10 118.046
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability