Jump to
S2C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -77.103461 |
| Energy at 298.15K | -77.102935 |
| HF Energy | -76.794083 |
| Nuclear repulsion energy | 23.983505 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3194 |
3032 |
57.11 |
|
|
|
| 2 |
A1 |
1708 |
1622 |
89.91 |
|
|
|
| 3 |
A1 |
1224 |
1162 |
21.72 |
|
|
|
| 4 |
B1 |
752 |
714 |
80.94 |
|
|
|
| 5 |
B2 |
3273 |
3107 |
27.86 |
|
|
|
| 6 |
B2 |
175 |
166 |
8.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5162.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 4901.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.478 |
| C2 |
0.000 |
0.000 |
0.815 |
| H3 |
0.000 |
0.939 |
-1.011 |
| H4 |
0.000 |
-0.939 |
-1.011 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.2937 | 1.0793 | 1.0793 |
C2 | 1.2937 | | 2.0535 | 2.0535 | H3 | 1.0793 | 2.0535 | | 1.8772 | H4 | 1.0793 | 2.0535 | 1.8772 | |
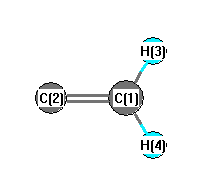 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
119.582 |
|
C2 |
C1 |
H4 |
119.582 |
| H3 |
C1 |
H4 |
120.836 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -77.023491 |
| Energy at 298.15K | -77.023624 |
| HF Energy | -76.758735 |
| Nuclear repulsion energy | 24.001294 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3154 |
2995 |
2.24 |
|
|
|
| 2 |
A1 |
1684 |
1599 |
2.29 |
|
|
|
| 3 |
A1 |
1444 |
1371 |
6.11 |
|
|
|
| 4 |
B1 |
996 |
946 |
32.12 |
|
|
|
| 5 |
B2 |
3221 |
3058 |
1.48 |
|
|
|
| 6 |
B2 |
1024 |
972 |
2.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5761.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 5470.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.473 |
| C2 |
0.000 |
0.000 |
0.817 |
| H3 |
0.000 |
0.926 |
-1.030 |
| H4 |
0.000 |
-0.926 |
-1.030 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.2903 | 1.0803 | 1.0803 |
C2 | 1.2903 | | 2.0662 | 2.0662 | H3 | 1.0803 | 2.0662 | | 1.8514 | H4 | 1.0803 | 2.0662 | 1.8514 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
