return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4N (pyrrolide radical)

using model chemistry: CCSD(T)/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at CCSD(T)/cc-pVTZ
 hartrees
Energy at 0K-209.131947
Energy at 298.15K-209.136608
HF Energy-208.266941
Nuclear repulsion energy150.665840
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3267 3184        
2 A1 3221 3140        
3 A1 1576 1537        
4 A1 1472 1435        
5 A1 1211 1180        
6 A1 1117 1089        
7 A1 1072 1045        
8 A1 876 854        
9 A2 894 872        
10 A2 837 816        
11 A2 498 485        
12 B1 836 815        
13 B1 720 702        
14 B1 547 533        
15 B2 3246 3164        
16 B2 3217 3136        
17 B2 1554 1515        
18 B2 1324 1291        
19 B2 1192 1162        
20 B2 1055 1028        
21 B2 801 781        

Unscaled Zero Point Vibrational Energy (zpe) 15265.9 cm-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 14881.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD(T)/cc-pVTZ
ABC
0.33247 0.29247 0.15560

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD(T)/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.251
C2 0.000 1.064 0.425
C3 0.000 -1.064 0.425
C4 0.000 0.682 -0.986
C5 0.000 -0.682 -0.986
H6 0.000 2.072 0.818
H7 0.000 -2.072 0.818
H8 0.000 1.354 -1.830
H9 0.000 -1.354 -1.830

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9
N11.34761.34762.33922.33922.11702.11703.36583.3658
C21.34762.12871.46182.24521.08173.16102.27343.3065
C31.34762.12872.24521.46183.16101.08173.30652.2734
C42.33921.46182.24521.36422.27743.29251.07872.2041
C52.33922.24521.46181.36423.29252.27742.20411.0787
H62.11701.08173.16102.27743.29254.14432.74364.3301
H72.11703.16101.08173.29252.27744.14434.33012.7436
H83.36582.27343.30651.07872.20412.74364.33012.7080
H93.36583.30652.27342.20411.07874.33012.74362.7080

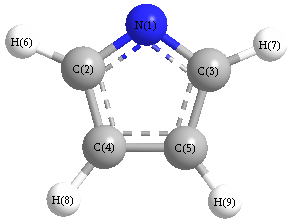
picture of pyrrolide radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 112.677 N1 C2 H6 120.861
N1 C3 C5 112.677 N1 C3 H7 120.861
C2 N1 C3 104.333 C2 C4 C5 105.157
C2 C4 H8 126.318 C3 C5 C4 105.157
C3 C5 H9 126.318 C4 C2 H6 126.463
C4 C5 H9 128.525 C5 C3 H7 126.463
C5 C4 H8 128.525
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability