return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHOH (1-hydroxy-ethyl radical)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-154.128278
Energy at 298.15K-154.133326
HF Energy-153.518559
Nuclear repulsion energy75.457631
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3888 3692 68.55      
2 A 3217 3055 36.67      
3 A 3181 3020 6.03      
4 A 3134 2975 12.35      
5 A 3052 2897 27.01      
6 A 1522 1445 2.74      
7 A 1498 1422 11.50      
8 A 1483 1408 19.94      
9 A 1407 1336 0.25      
10 A 1293 1228 118.09      
11 A 1239 1177 44.87      
12 A 1080 1025 28.22      
13 A 1044 991 5.30      
14 A 952 904 11.32      
15 A 624 593 16.58      
16 A 417 396 9.39      
17 A 379 360 120.33      
18 A 198 188 4.08      

Unscaled Zero Point Vibrational Energy (zpe) 14804.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14055.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.54532 0.32065 0.28093

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.090 0.509 -0.105
C2 1.217 -0.165 0.013
O3 -1.157 -0.341 0.021
H4 -0.227 1.513 0.259
H5 1.270 -1.006 -0.671
H6 2.020 0.524 -0.225
H7 1.389 -0.550 1.020
H8 -1.963 0.179 0.006

Atom - Atom Distances (Å)
  C1 C2 O3 H4 H5 H6 H7 H8
C11.47491.36961.07712.11242.11302.13791.9054
C21.47492.38092.22701.08561.08391.09143.1990
O31.36962.38092.08762.60953.30152.74290.9592
H41.07712.22702.08763.07402.50182.72822.2043
H52.11241.08562.60953.07401.76141.75513.5090
H62.11301.08393.30152.50181.76141.76104.0045
H72.13791.09142.74292.72821.75511.76103.5770
H81.90543.19900.95922.20433.50904.00453.5770

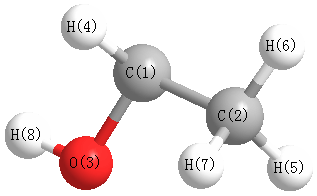
picture of 1-hydroxy-ethyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 110.251 C1 C2 H6 110.401
C1 C2 H7 111.967 C1 O3 H8 108.523
C2 C1 O3 113.600 C2 C1 H4 120.743
O3 C1 H4 116.624 H5 C2 H6 108.559
H5 C2 H7 107.448 H6 C2 H7 108.098
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability