Jump to
S1C2
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.372132 |
| Energy at 298.15K | -100.371210 |
| HF Energy | -100.372132 |
| Nuclear repulsion energy | 27.800313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.057 |
| C2 |
0.000 |
0.000 |
-0.149 |
| N3 |
0.000 |
0.000 |
1.009 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9079 | 3.0665 |
C2 | 1.9079 | | 1.1585 | N3 | 3.0665 | 1.1585 | |
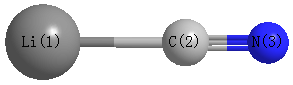 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.399 |
|
|
|
| 2 |
C |
-0.251 |
|
|
|
| 3 |
N |
-0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-9.021 |
9.021 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.171 |
0.000 |
0.000 |
| y |
0.000 |
-14.171 |
0.000 |
| z |
0.000 |
0.000 |
1.171 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.671 |
0.000 |
0.000 |
| y |
0.000 |
-7.671 |
0.000 |
| z |
0.000 |
0.000 |
15.342 |
|
| Polar |
|---|
| 3z2-r2 | 30.685 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.686 |
0.000 |
0.000 |
| y |
0.000 |
2.686 |
0.000 |
| z |
0.000 |
0.000 |
4.252 |
<r2> (average value of r
2) Å
2
| <r2> |
25.617 |
| (<r2>)1/2 |
5.061 |
Jump to
S1C1
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.375788 |
| Energy at 298.15K | |
| HF Energy | -100.375788 |
| Nuclear repulsion energy | 29.401211 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.506 |
-0.483 |
0.000 |
| C2 |
-0.753 |
-0.368 |
0.000 |
| N3 |
0.000 |
0.522 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.2625 | 1.8111 |
C2 | 2.2625 | | 1.1664 | N3 | 1.8111 | 1.1664 | |
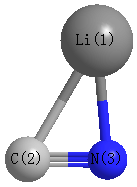 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
52.687 |
|
Li1 |
N3 |
C2 |
96.503 |
| C2 |
Li1 |
N3 |
30.810 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.308 |
|
|
|
| 2 |
C |
-0.155 |
|
|
|
| 3 |
N |
-0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.924 |
-1.989 |
0.000 |
7.204 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-5.226 |
-5.455 |
0.000 |
| y |
-5.455 |
-15.304 |
0.000 |
| z |
0.000 |
0.000 |
-14.311 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.582 |
-5.455 |
0.000 |
| y |
-5.455 |
-5.536 |
0.000 |
| z |
0.000 |
0.000 |
-4.046 |
|
| Polar |
|---|
| 3z2-r2 | -8.092 |
|---|
| x2-y2 | 10.079 |
|---|
| xy | -5.455 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.686 |
0.000 |
0.000 |
| y |
0.000 |
2.686 |
0.000 |
| z |
0.000 |
0.000 |
4.252 |
<r2> (average value of r
2) Å
2
| <r2> |
20.889 |
| (<r2>)1/2 |
4.570 |
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.376807 |
| Energy at 298.15K | -100.375707 |
| HF Energy | -100.376807 |
| Nuclear repulsion energy | 28.549139 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.869 |
| C2 |
0.000 |
0.000 |
-1.062 |
| N3 |
0.000 |
0.000 |
0.109 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9306 | 1.7599 |
C2 | 2.9306 | | 1.1707 | N3 | 1.7599 | 1.1707 | |
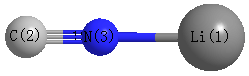 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.369 |
|
|
|
| 2 |
C |
-0.290 |
|
|
|
| 3 |
N |
-0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
8.821 |
8.821 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.209 |
0.000 |
0.000 |
| y |
0.000 |
-14.209 |
0.000 |
| z |
0.000 |
0.000 |
-2.469 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.870 |
0.000 |
0.000 |
| y |
0.000 |
-5.870 |
0.000 |
| z |
0.000 |
0.000 |
11.740 |
|
| Polar |
|---|
| 3z2-r2 | 23.480 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.773 |
0.000 |
0.000 |
| y |
0.000 |
2.773 |
0.000 |
| z |
0.000 |
0.000 |
4.416 |
<r2> (average value of r
2) Å
2
| <r2> |
23.755 |
| (<r2>)1/2 |
4.874 |
