Jump to
S1C2
S2C1
S2C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -39.090397 |
| Energy at 298.15K | -39.090211 |
| HF Energy | -38.937802 |
| Nuclear repulsion energy | 6.193125 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.105 |
| H2 |
0.000 |
0.986 |
-0.316 |
| H3 |
0.000 |
-0.986 |
-0.316 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0718 | 1.0718 |
H2 | 1.0718 | | 1.9711 | H3 | 1.0718 | 1.9711 | |
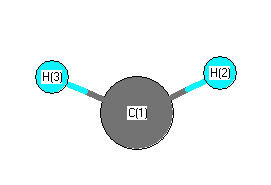 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
133.715 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -39.080787 |
| Energy at 298.15K | |
| HF Energy | -38.925883 |
| Nuclear repulsion energy | 6.227544 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| H2 |
0.000 |
0.000 |
1.062 |
| H3 |
0.000 |
0.000 |
-1.062 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0616 | 1.0616 |
H2 | 1.0616 | | 2.1233 | H3 | 1.0616 | 2.1233 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -39.070759 |
| Energy at 298.15K | -39.070596 |
| HF Energy | -38.892397 |
| Nuclear repulsion energy | 6.057595 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.174 |
| H2 |
0.000 |
0.857 |
-0.523 |
| H3 |
0.000 |
-0.857 |
-0.523 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.1046 | 1.1046 |
H2 | 1.1046 | | 1.7139 | H3 | 1.1046 | 1.7139 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -39.070759 |
| Energy at 298.15K | -39.070596 |
| HF Energy | -38.892397 |
| Nuclear repulsion energy | 6.057595 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
