return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-307.815844
Energy at 298.15K-307.821571
HF Energy-306.443021
Nuclear repulsion energy316.262442
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3291 3125 0.00      
2 Ag 3267 3101 0.00      
3 Ag 3251 3087 0.00      
4 Ag 1632 1549 0.00      
5 Ag 1544 1466 0.00      
6 Ag 1402 1331 0.00      
7 Ag 1330 1263 0.00      
8 Ag 1203 1142 0.00      
9 Ag 1114 1058 0.00      
10 Ag 979 929 0.00      
11 Ag 867 823 0.00      
12 Ag 717 681 0.00      
13 Ag 662 629 0.00      
14 Au 922 875 8.52      
15 Au 884 839 14.67      
16 Au 731 694 93.37      
17 Au 511 485 9.30      
18 Au 298 283 1.44      
19 Au 154 146 6.63      
20 Bg 927 881 0.00      
21 Bg 914 868 0.00      
22 Bg 783 744 0.00      
23 Bg 667 633 0.00      
24 Bg 552 524 0.00      
25 Bu 3291 3124 16.65      
26 Bu 3267 3102 5.57      
27 Bu 3252 3087 10.11      
28 Bu 1637 1555 41.32      
29 Bu 1524 1447 12.15      
30 Bu 1349 1281 16.70      
31 Bu 1278 1214 16.80      
32 Bu 1138 1080 39.66      
33 Bu 1068 1014 3.17      
34 Bu 968 919 6.28      
35 Bu 846 804 5.32      
36 Bu 439 417 3.41      

Unscaled Zero Point Vibrational Energy (zpe) 24329.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23098.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.17681 0.07301 0.05167

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.336 1.088 0.000
C2 2.154 -0.157 0.000
C3 1.336 -1.232 0.000
C4 -1.336 -1.088 0.000
C5 -2.154 0.157 0.000
C6 -1.336 1.232 0.000
C7 0.038 0.725 0.000
C8 -0.038 -0.725 0.000
H9 1.752 2.080 0.000
H10 3.228 -0.170 0.000
H11 1.632 -2.264 0.000
H12 -1.752 -2.080 0.000
H13 -3.228 0.170 0.000
H14 -1.632 2.264 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.48932.31953.44603.61202.67631.34822.27481.07552.27163.36454.42364.65573.1928
C21.48931.35033.61204.31913.75632.29282.26402.27271.07432.17034.35325.39184.4938
C32.31951.35032.67633.75633.63472.34871.46423.33752.16961.07363.20224.77454.5858
C43.44603.61202.67631.48932.31952.27481.34824.42364.65573.19281.07552.27163.3645
C53.61204.31913.75631.48931.35032.26402.29284.35325.39184.49382.27271.07432.1703
C62.67633.75633.63472.31951.35031.46422.34873.20224.77454.58583.33752.16961.0736
C71.34822.29282.34872.27482.26401.46421.45262.18463.31353.38773.32723.31262.2704
C82.27482.26401.46421.34822.29282.34871.45263.32723.31262.27042.18463.31353.3877
H91.07552.27273.33754.42364.35323.20222.18463.32722.69064.34515.43835.33353.3888
H102.27161.07432.16964.65575.39184.77453.31353.31262.69062.63295.33356.46505.4353
H113.36452.17031.07363.19284.49384.58583.38772.27044.34512.63293.38885.43535.5815
H124.42364.35323.20221.07552.27273.33753.32722.18465.43835.33353.38882.69064.3451
H134.65575.39184.77452.27161.07432.16963.31263.31355.33356.46505.43532.69062.6329
H143.19284.49384.58583.36452.17031.07362.27043.38773.38885.43535.58154.34512.6329

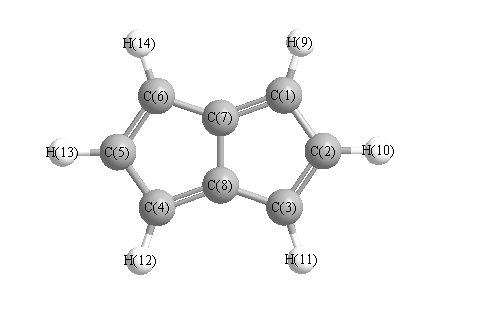
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.439 C1 C2 H10 123.965
C1 C7 C6 144.172 C1 C7 C8 108.565
C2 C1 C7 107.703 C2 C1 H9 123.979
C2 C3 C8 107.031 C2 C3 H11 126.736
C3 C2 H10 126.596 C3 C8 C4 144.172
C3 C8 C7 107.263 C4 C5 C6 109.439
C4 C5 H13 123.965 C4 C8 C7 108.565
C5 C4 C8 107.703 C5 C4 H12 123.979
C5 C6 C7 107.031 C5 C6 H14 126.736
C6 C5 H13 126.596 C6 C7 C8 107.263
C7 C1 H9 128.318 C7 C6 H14 126.233
C8 C3 H11 126.233 C8 C4 H12 128.318
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability