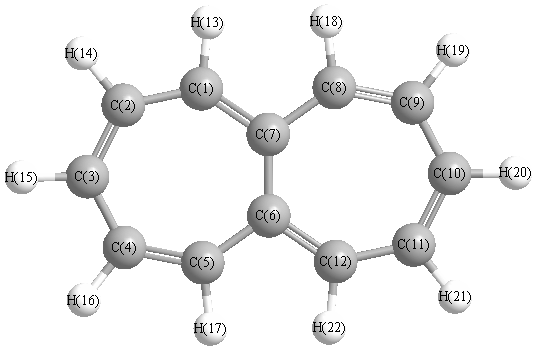
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3198 |
3061 |
3.25 |
|
|
|
| 2 |
A |
3179 |
3042 |
18.47 |
|
|
|
| 3 |
A |
3174 |
3037 |
0.57 |
|
|
|
| 4 |
A |
3157 |
3021 |
0.02 |
|
|
|
| 5 |
A |
3156 |
3020 |
0.62 |
|
|
|
| 6 |
A |
1722 |
1648 |
3.24 |
|
|
|
| 7 |
A |
1669 |
1597 |
3.58 |
|
|
|
| 8 |
A |
1625 |
1555 |
0.71 |
|
|
|
| 9 |
A |
1487 |
1423 |
0.01 |
|
|
|
| 10 |
A |
1450 |
1388 |
0.05 |
|
|
|
| 11 |
A |
1417 |
1356 |
1.03 |
|
|
|
| 12 |
A |
1270 |
1216 |
0.02 |
|
|
|
| 13 |
A |
1252 |
1198 |
0.05 |
|
|
|
| 14 |
A |
1217 |
1165 |
0.34 |
|
|
|
| 15 |
A |
1073 |
1027 |
1.05 |
|
|
|
| 16 |
A |
1015 |
972 |
0.67 |
|
|
|
| 17 |
A |
1007 |
963 |
0.30 |
|
|
|
| 18 |
A |
977 |
935 |
0.27 |
|
|
|
| 19 |
A |
916 |
876 |
13.14 |
|
|
|
| 20 |
A |
896 |
857 |
0.97 |
|
|
|
| 21 |
A |
831 |
796 |
3.44 |
|
|
|
| 22 |
A |
786 |
753 |
0.67 |
|
|
|
| 23 |
A |
730 |
698 |
65.87 |
|
|
|
| 24 |
A |
669 |
640 |
2.87 |
|
|
|
| 25 |
A |
585 |
560 |
0.01 |
|
|
|
| 26 |
A |
426 |
408 |
5.58 |
|
|
|
| 27 |
A |
382 |
366 |
5.67 |
|
|
|
| 28 |
A |
353 |
338 |
0.46 |
|
|
|
| 29 |
A |
289 |
277 |
1.76 |
|
|
|
| 30 |
A |
128 |
122 |
1.08 |
|
|
|
| 31 |
A |
67 |
64 |
0.01 |
|
|
|
| 32 |
B |
3197 |
3060 |
59.48 |
|
|
|
| 33 |
B |
3179 |
3043 |
45.38 |
|
|
|
| 34 |
B |
3172 |
3036 |
3.78 |
|
|
|
| 35 |
B |
3158 |
3023 |
3.82 |
|
|
|
| 36 |
B |
3155 |
3020 |
16.43 |
|
|
|
| 37 |
B |
1691 |
1618 |
19.68 |
|
|
|
| 38 |
B |
1669 |
1597 |
66.77 |
|
|
|
| 39 |
B |
1594 |
1525 |
18.95 |
|
|
|
| 40 |
B |
1486 |
1422 |
2.91 |
|
|
|
| 41 |
B |
1416 |
1355 |
4.22 |
|
|
|
| 42 |
B |
1336 |
1279 |
1.36 |
|
|
|
| 43 |
B |
1266 |
1211 |
7.20 |
|
|
|
| 44 |
B |
1247 |
1194 |
0.96 |
|
|
|
| 45 |
B |
1142 |
1093 |
1.87 |
|
|
|
| 46 |
B |
1050 |
1005 |
2.44 |
|
|
|
| 47 |
B |
1013 |
969 |
0.67 |
|
|
|
| 48 |
B |
1004 |
961 |
2.28 |
|
|
|
| 49 |
B |
954 |
913 |
4.08 |
|
|
|
| 50 |
B |
933 |
893 |
0.85 |
|
|
|
| 51 |
B |
880 |
842 |
12.87 |
|
|
|
| 52 |
B |
819 |
784 |
50.43 |
|
|
|
| 53 |
B |
753 |
721 |
15.49 |
|
|
|
| 54 |
B |
664 |
635 |
9.59 |
|
|
|
| 55 |
B |
600 |
574 |
5.84 |
|
|
|
| 56 |
B |
583 |
558 |
17.29 |
|
|
|
| 57 |
B |
413 |
395 |
1.19 |
|
|
|
| 58 |
B |
354 |
338 |
1.09 |
|
|
|
| 59 |
B |
279 |
267 |
0.44 |
|
|
|
| 60 |
B |
155 |
148 |
2.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39630.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 37929.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.200 |
|
|
|
| 2 |
C |
-0.121 |
|
|
|
| 3 |
C |
-0.149 |
|
|
|
| 4 |
C |
-0.095 |
|
|
|
| 5 |
C |
-0.198 |
|
|
|
| 6 |
C |
0.130 |
|
|
|
| 7 |
C |
0.130 |
|
|
|
| 8 |
C |
-0.198 |
|
|
|
| 9 |
C |
-0.095 |
|
|
|
| 10 |
C |
-0.149 |
|
|
|
| 11 |
C |
-0.121 |
|
|
|
| 12 |
C |
-0.200 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.124 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.142 |
|
|
|
| 18 |
H |
0.142 |
|
|
|
| 19 |
H |
0.128 |
|
|
|
| 20 |
H |
0.125 |
|
|
|
| 21 |
H |
0.124 |
|
|
|
| 22 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.233 |
0.233 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-64.207 |
-0.886 |
0.000 |
| y |
-0.886 |
-62.684 |
0.000 |
| z |
0.000 |
0.000 |
-72.630 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.450 |
-0.886 |
0.000 |
| y |
-0.886 |
5.734 |
0.000 |
| z |
0.000 |
0.000 |
-9.184 |
|
| Polar |
|---|
| 3z2-r2 | -18.368 |
|---|
| x2-y2 | -1.523 |
|---|
| xy | -0.886 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
20.206 |
0.070 |
0.000 |
| y |
0.070 |
31.185 |
0.000 |
| z |
0.000 |
0.000 |
12.038 |
<r2> (average value of r
2) Å
2
| <r2> |
539.036 |
| (<r2>)1/2 |
23.217 |