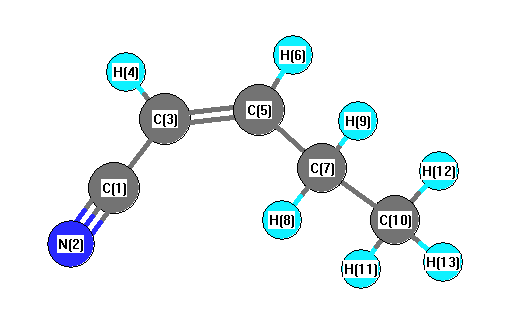
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3211 |
3073 |
2.51 |
|
|
|
| 2 |
A |
3172 |
3036 |
8.56 |
|
|
|
| 3 |
A |
3144 |
3009 |
20.77 |
|
|
|
| 4 |
A |
3136 |
3002 |
23.09 |
|
|
|
| 5 |
A |
3107 |
2974 |
0.69 |
|
|
|
| 6 |
A |
3061 |
2930 |
26.91 |
|
|
|
| 7 |
A |
3045 |
2915 |
10.55 |
|
|
|
| 8 |
A |
2363 |
2262 |
19.10 |
|
|
|
| 9 |
A |
1709 |
1636 |
19.09 |
|
|
|
| 10 |
A |
1506 |
1442 |
7.61 |
|
|
|
| 11 |
A |
1499 |
1435 |
6.66 |
|
|
|
| 12 |
A |
1482 |
1418 |
5.27 |
|
|
|
| 13 |
A |
1427 |
1366 |
0.87 |
|
|
|
| 14 |
A |
1407 |
1347 |
1.13 |
|
|
|
| 15 |
A |
1342 |
1285 |
1.95 |
|
|
|
| 16 |
A |
1296 |
1240 |
0.07 |
|
|
|
| 17 |
A |
1252 |
1199 |
0.71 |
|
|
|
| 18 |
A |
1146 |
1097 |
0.46 |
|
|
|
| 19 |
A |
1094 |
1047 |
3.21 |
|
|
|
| 20 |
A |
1046 |
1001 |
4.31 |
|
|
|
| 21 |
A |
1010 |
967 |
0.95 |
|
|
|
| 22 |
A |
972 |
930 |
4.55 |
|
|
|
| 23 |
A |
876 |
838 |
3.98 |
|
|
|
| 24 |
A |
806 |
771 |
7.26 |
|
|
|
| 25 |
A |
769 |
736 |
34.22 |
|
|
|
| 26 |
A |
676 |
647 |
1.89 |
|
|
|
| 27 |
A |
577 |
552 |
1.15 |
|
|
|
| 28 |
A |
401 |
384 |
0.35 |
|
|
|
| 29 |
A |
364 |
348 |
0.02 |
|
|
|
| 30 |
A |
241 |
231 |
0.69 |
|
|
|
| 31 |
A |
214 |
205 |
2.92 |
|
|
|
| 32 |
A |
146 |
139 |
4.75 |
|
|
|
| 33 |
A |
61 |
58 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23779.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22759.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
N |
-0.098 |
|
|
|
| 3 |
C |
-0.147 |
|
|
|
| 4 |
H |
0.164 |
|
|
|
| 5 |
C |
-0.107 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
C |
-0.198 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
| 9 |
H |
0.130 |
|
|
|
| 10 |
C |
-0.349 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.113 |
|
|
|
| 13 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.413 |
2.484 |
0.074 |
4.222 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.498 |
-5.600 |
1.033 |
| y |
-5.600 |
-35.932 |
-0.676 |
| z |
1.033 |
-0.676 |
-36.938 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.063 |
-5.600 |
1.033 |
| y |
-5.600 |
3.786 |
-0.676 |
| z |
1.033 |
-0.676 |
2.277 |
|
| Polar |
|---|
| 3z2-r2 | 4.553 |
|---|
| x2-y2 | -6.566 |
|---|
| xy | -5.600 |
|---|
| xz | 1.033 |
|---|
| yz | -0.676 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.474 |
0.899 |
0.426 |
| y |
0.899 |
8.889 |
-0.218 |
| z |
0.426 |
-0.218 |
6.605 |
<r2> (average value of r
2) Å
2
| <r2> |
198.400 |
| (<r2>)1/2 |
14.085 |