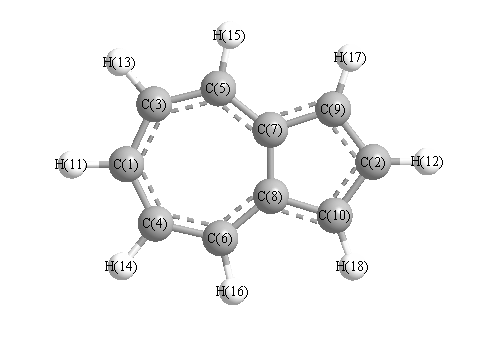
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3253 |
3113 |
9.59 |
|
|
|
| 2 |
A1 |
3224 |
3086 |
6.84 |
|
|
|
| 3 |
A1 |
3199 |
3062 |
18.72 |
|
|
|
| 4 |
A1 |
3170 |
3034 |
6.45 |
|
|
|
| 5 |
A1 |
3160 |
3024 |
0.08 |
|
|
|
| 6 |
A1 |
1667 |
1595 |
67.55 |
|
|
|
| 7 |
A1 |
1598 |
1530 |
8.47 |
|
|
|
| 8 |
A1 |
1503 |
1438 |
32.77 |
|
|
|
| 9 |
A1 |
1439 |
1378 |
84.31 |
|
|
|
| 10 |
A1 |
1317 |
1260 |
1.10 |
|
|
|
| 11 |
A1 |
1245 |
1192 |
0.12 |
|
|
|
| 12 |
A1 |
1089 |
1042 |
7.00 |
|
|
|
| 13 |
A1 |
973 |
932 |
2.24 |
|
|
|
| 14 |
A1 |
911 |
872 |
2.64 |
|
|
|
| 15 |
A1 |
833 |
797 |
6.00 |
|
|
|
| 16 |
A1 |
690 |
660 |
0.85 |
|
|
|
| 17 |
A1 |
404 |
386 |
1.43 |
|
|
|
| 18 |
A2 |
1006 |
963 |
0.00 |
|
|
|
| 19 |
A2 |
887 |
849 |
0.00 |
|
|
|
| 20 |
A2 |
807 |
772 |
0.00 |
|
|
|
| 21 |
A2 |
733 |
702 |
0.00 |
|
|
|
| 22 |
A2 |
432 |
414 |
0.00 |
|
|
|
| 23 |
A2 |
161 |
154 |
0.00 |
|
|
|
| 24 |
B1 |
1023 |
979 |
0.04 |
|
|
|
| 25 |
B1 |
988 |
946 |
5.62 |
|
|
|
| 26 |
B1 |
952 |
911 |
0.04 |
|
|
|
| 27 |
B1 |
794 |
760 |
108.28 |
|
|
|
| 28 |
B1 |
748 |
716 |
4.49 |
|
|
|
| 29 |
B1 |
618 |
591 |
1.23 |
|
|
|
| 30 |
B1 |
576 |
551 |
9.30 |
|
|
|
| 31 |
B1 |
320 |
306 |
9.76 |
|
|
|
| 32 |
B1 |
169 |
162 |
1.81 |
|
|
|
| 33 |
B2 |
3244 |
3105 |
11.26 |
|
|
|
| 34 |
B2 |
3191 |
3054 |
27.63 |
|
|
|
| 35 |
B2 |
3162 |
3026 |
12.54 |
|
|
|
| 36 |
B2 |
1666 |
1595 |
4.77 |
|
|
|
| 37 |
B2 |
1559 |
1492 |
8.70 |
|
|
|
| 38 |
B2 |
1497 |
1432 |
7.82 |
|
|
|
| 39 |
B2 |
1437 |
1375 |
0.62 |
|
|
|
| 40 |
B2 |
1338 |
1281 |
1.19 |
|
|
|
| 41 |
B2 |
1321 |
1264 |
3.47 |
|
|
|
| 42 |
B2 |
1241 |
1188 |
7.10 |
|
|
|
| 43 |
B2 |
1191 |
1140 |
0.29 |
|
|
|
| 44 |
B2 |
1068 |
1022 |
0.32 |
|
|
|
| 45 |
B2 |
1023 |
979 |
13.94 |
|
|
|
| 46 |
B2 |
742 |
710 |
0.49 |
|
|
|
| 47 |
B2 |
494 |
473 |
1.77 |
|
|
|
| 48 |
B2 |
326 |
312 |
0.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32193.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30812.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.089 |
|
|
|
| 2 |
C |
-0.124 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.156 |
|
|
|
| 5 |
C |
-0.095 |
|
|
|
| 6 |
C |
-0.095 |
|
|
|
| 7 |
C |
0.120 |
|
|
|
| 8 |
C |
0.120 |
|
|
|
| 9 |
C |
-0.268 |
|
|
|
| 10 |
C |
-0.268 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.125 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.060 |
1.060 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-63.314 |
0.000 |
0.000 |
| y |
0.000 |
-50.679 |
0.000 |
| z |
0.000 |
0.000 |
-51.510 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.219 |
0.000 |
0.000 |
| y |
0.000 |
6.733 |
0.000 |
| z |
0.000 |
0.000 |
5.486 |
|
| Polar |
|---|
| 3z2-r2 | 10.972 |
|---|
| x2-y2 | -12.635 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.655 |
0.000 |
0.000 |
| y |
0.000 |
18.318 |
0.000 |
| z |
0.000 |
0.000 |
27.346 |
<r2> (average value of r
2) Å
2
| <r2> |
361.216 |
| (<r2>)1/2 |
19.006 |