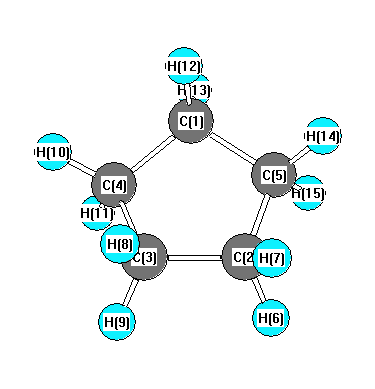
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3121 |
2987 |
95.02 |
|
|
|
| 2 |
A |
3106 |
2972 |
22.50 |
|
|
|
| 3 |
A |
3104 |
2971 |
28.91 |
|
|
|
| 4 |
A |
3101 |
2968 |
13.07 |
|
|
|
| 5 |
A |
3090 |
2958 |
34.86 |
|
|
|
| 6 |
A |
3071 |
2939 |
52.58 |
|
|
|
| 7 |
A |
3056 |
2925 |
23.54 |
|
|
|
| 8 |
A |
3051 |
2920 |
29.20 |
|
|
|
| 9 |
A |
3044 |
2913 |
35.03 |
|
|
|
| 10 |
A |
3037 |
2907 |
17.60 |
|
|
|
| 11 |
A |
1516 |
1451 |
0.20 |
|
|
|
| 12 |
A |
1490 |
1426 |
4.70 |
|
|
|
| 13 |
A |
1489 |
1425 |
3.13 |
|
|
|
| 14 |
A |
1484 |
1420 |
4.59 |
|
|
|
| 15 |
A |
1479 |
1416 |
0.22 |
|
|
|
| 16 |
A |
1347 |
1289 |
0.05 |
|
|
|
| 17 |
A |
1347 |
1289 |
0.09 |
|
|
|
| 18 |
A |
1338 |
1280 |
0.02 |
|
|
|
| 19 |
A |
1320 |
1263 |
0.59 |
|
|
|
| 20 |
A |
1302 |
1246 |
0.03 |
|
|
|
| 21 |
A |
1284 |
1228 |
0.84 |
|
|
|
| 22 |
A |
1235 |
1182 |
0.15 |
|
|
|
| 23 |
A |
1224 |
1172 |
0.28 |
|
|
|
| 24 |
A |
1202 |
1151 |
0.18 |
|
|
|
| 25 |
A |
1184 |
1133 |
0.01 |
|
|
|
| 26 |
A |
1065 |
1020 |
0.04 |
|
|
|
| 27 |
A |
1049 |
1004 |
0.00 |
|
|
|
| 28 |
A |
996 |
953 |
0.55 |
|
|
|
| 29 |
A |
964 |
923 |
1.41 |
|
|
|
| 30 |
A |
928 |
888 |
1.75 |
|
|
|
| 31 |
A |
920 |
880 |
1.80 |
|
|
|
| 32 |
A |
904 |
865 |
0.76 |
|
|
|
| 33 |
A |
874 |
837 |
0.81 |
|
|
|
| 34 |
A |
824 |
789 |
0.20 |
|
|
|
| 35 |
A |
764 |
732 |
0.73 |
|
|
|
| 36 |
A |
611 |
585 |
0.09 |
|
|
|
| 37 |
A |
536 |
513 |
1.20 |
|
|
|
| 38 |
A |
265 |
254 |
0.01 |
|
|
|
| 39 |
A |
27 |
26 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30873.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29548.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.208 |
|
|
|
| 2 |
C |
-0.220 |
|
|
|
| 3 |
C |
-0.220 |
|
|
|
| 4 |
C |
-0.217 |
|
|
|
| 5 |
C |
-0.210 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.107 |
|
|
|
| 11 |
H |
0.108 |
|
|
|
| 12 |
H |
0.107 |
|
|
|
| 13 |
H |
0.105 |
|
|
|
| 14 |
H |
0.105 |
|
|
|
| 15 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.020 |
-0.008 |
0.001 |
0.021 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.767 |
0.006 |
0.006 |
| y |
0.006 |
-33.752 |
-0.038 |
| z |
0.006 |
-0.038 |
-32.521 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.630 |
0.006 |
0.006 |
| y |
0.006 |
-0.608 |
-0.038 |
| z |
0.006 |
-0.038 |
1.238 |
|
| Polar |
|---|
| 3z2-r2 | 2.476 |
|---|
| x2-y2 | -0.015 |
|---|
| xy | 0.006 |
|---|
| xz | 0.006 |
|---|
| yz | -0.038 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.836 |
0.004 |
-0.002 |
| y |
0.004 |
8.840 |
0.015 |
| z |
-0.002 |
0.015 |
7.724 |
<r2> (average value of r
2) Å
2
| <r2> |
114.318 |
| (<r2>)1/2 |
10.692 |