return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H3NO (Oxazole)

using model chemistry: QCISD(T)/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVTZ
 hartrees
Energy at 0K-245.677106
Energy at 298.15K-245.681716
HF Energy-244.718120
Nuclear repulsion energy162.865871
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3311 3141        
2 A' 3288 3119        
3 A' 3278 3109        
4 A' 1572 1492        
5 A' 1537 1458        
6 A' 1357 1287        
7 A' 1278 1212        
8 A' 1174 1114        
9 A' 1112 1055        
10 A' 1105 1048        
11 A' 1072 1017        
12 A' 917 870        
13 A' 908 861        
14 A" 871 826        
15 A" 849 806        
16 A" 761 722        
17 A" 655 621        
18 A" 618 586        

Unscaled Zero Point Vibrational Energy (zpe) 12831.5 cm-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 12172.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVTZ
ABC
0.33581 0.32038 0.16396

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.106 0.312 0.000
C2 0.000 1.099 0.000
N3 1.130 0.466 0.000
C4 0.754 -0.881 0.000
C5 -0.599 -0.965 0.000
H6 -0.165 2.163 0.000
H7 1.482 -1.674 0.000
H8 -1.312 -1.768 0.000

Atom - Atom Distances (Å)
  O1 C2 N3 C4 C5 H6 H7 H8
O11.35792.24202.21011.37342.07683.26192.0900
C21.35791.29552.11892.14891.07653.14403.1532
N32.24201.29551.39872.24462.13452.16873.3104
C42.21012.11891.39871.35603.17971.07592.2489
C51.37342.14892.24461.35603.15752.19831.0744
H62.07681.07652.13453.17973.15754.17514.0951
H73.26193.14402.16871.07592.19834.17512.7955
H82.09003.15323.31042.24891.07444.09512.7955

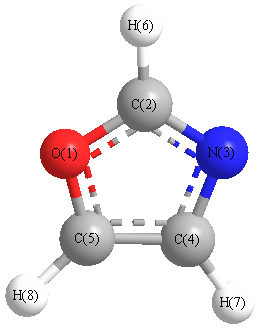
picture of Oxazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 N3 115.316 O1 C2 H6 116.632
O1 C5 C4 108.141 O1 C5 H8 116.740
C2 O1 C5 103.766 C2 N3 C4 103.650
N3 C2 H6 128.051 N3 C4 C5 109.127
N3 C4 H7 121.873 C4 C5 H8 135.119
C5 C4 H7 129.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability