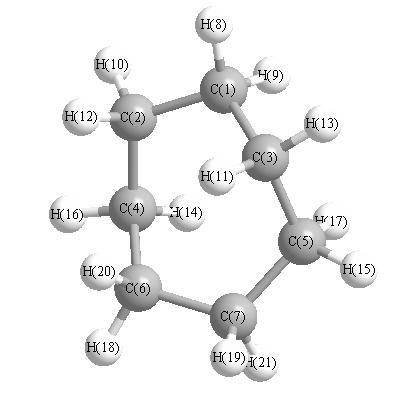
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3089 |
2957 |
5.11 |
|
|
|
| 2 |
A |
3077 |
2945 |
51.85 |
|
|
|
| 3 |
A |
3072 |
2940 |
64.63 |
|
|
|
| 4 |
A |
3046 |
2915 |
20.41 |
|
|
|
| 5 |
A |
3038 |
2907 |
0.50 |
|
|
|
| 6 |
A |
3036 |
2906 |
10.71 |
|
|
|
| 7 |
A |
3026 |
2896 |
11.60 |
|
|
|
| 8 |
A |
1511 |
1446 |
0.51 |
|
|
|
| 9 |
A |
1494 |
1430 |
3.72 |
|
|
|
| 10 |
A |
1488 |
1424 |
2.89 |
|
|
|
| 11 |
A |
1483 |
1420 |
0.39 |
|
|
|
| 12 |
A |
1397 |
1337 |
0.16 |
|
|
|
| 13 |
A |
1392 |
1333 |
0.79 |
|
|
|
| 14 |
A |
1373 |
1314 |
0.45 |
|
|
|
| 15 |
A |
1316 |
1259 |
0.14 |
|
|
|
| 16 |
A |
1306 |
1250 |
1.63 |
|
|
|
| 17 |
A |
1230 |
1177 |
0.56 |
|
|
|
| 18 |
A |
1217 |
1165 |
0.01 |
|
|
|
| 19 |
A |
1135 |
1086 |
0.10 |
|
|
|
| 20 |
A |
1057 |
1011 |
0.41 |
|
|
|
| 21 |
A |
1032 |
988 |
0.03 |
|
|
|
| 22 |
A |
968 |
927 |
1.51 |
|
|
|
| 23 |
A |
857 |
820 |
0.06 |
|
|
|
| 24 |
A |
846 |
810 |
2.11 |
|
|
|
| 25 |
A |
752 |
720 |
0.04 |
|
|
|
| 26 |
A |
513 |
491 |
0.15 |
|
|
|
| 27 |
A |
396 |
379 |
0.00 |
|
|
|
| 28 |
A |
311 |
298 |
0.00 |
|
|
|
| 29 |
A |
182 |
174 |
0.00 |
|
|
|
| 30 |
B |
3093 |
2961 |
107.91 |
|
|
|
| 31 |
B |
3084 |
2952 |
127.86 |
|
|
|
| 32 |
B |
3078 |
2946 |
35.32 |
|
|
|
| 33 |
B |
3069 |
2937 |
10.55 |
|
|
|
| 34 |
B |
3040 |
2910 |
54.16 |
|
|
|
| 35 |
B |
3030 |
2900 |
8.25 |
|
|
|
| 36 |
B |
3028 |
2898 |
26.26 |
|
|
|
| 37 |
B |
1507 |
1443 |
20.77 |
|
|
|
| 38 |
B |
1490 |
1426 |
9.41 |
|
|
|
| 39 |
B |
1479 |
1416 |
1.33 |
|
|
|
| 40 |
B |
1396 |
1336 |
0.90 |
|
|
|
| 41 |
B |
1384 |
1324 |
0.03 |
|
|
|
| 42 |
B |
1379 |
1320 |
0.51 |
|
|
|
| 43 |
B |
1352 |
1294 |
0.18 |
|
|
|
| 44 |
B |
1307 |
1251 |
0.94 |
|
|
|
| 45 |
B |
1225 |
1172 |
0.45 |
|
|
|
| 46 |
B |
1209 |
1157 |
0.91 |
|
|
|
| 47 |
B |
1147 |
1098 |
0.39 |
|
|
|
| 48 |
B |
1036 |
992 |
0.30 |
|
|
|
| 49 |
B |
1025 |
981 |
0.70 |
|
|
|
| 50 |
B |
971 |
929 |
2.23 |
|
|
|
| 51 |
B |
873 |
836 |
0.36 |
|
|
|
| 52 |
B |
822 |
787 |
3.13 |
|
|
|
| 53 |
B |
736 |
705 |
1.55 |
|
|
|
| 54 |
B |
486 |
465 |
0.30 |
|
|
|
| 55 |
B |
337 |
323 |
0.04 |
|
|
|
| 56 |
B |
270 |
258 |
0.07 |
|
|
|
| 57 |
B |
146 |
139 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43819.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 41939.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.235 |
|
|
|
| 2 |
C |
-0.237 |
|
|
|
| 3 |
C |
-0.237 |
|
|
|
| 4 |
C |
-0.234 |
|
|
|
| 5 |
C |
-0.234 |
|
|
|
| 6 |
C |
-0.236 |
|
|
|
| 7 |
C |
-0.236 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.118 |
|
|
|
| 12 |
H |
0.118 |
|
|
|
| 13 |
H |
0.118 |
|
|
|
| 14 |
H |
0.120 |
|
|
|
| 15 |
H |
0.120 |
|
|
|
| 16 |
H |
0.117 |
|
|
|
| 17 |
H |
0.117 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
| 19 |
H |
0.119 |
|
|
|
| 20 |
H |
0.117 |
|
|
|
| 21 |
H |
0.117 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.013 |
0.013 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.493 |
-0.136 |
0.000 |
| y |
-0.136 |
-46.459 |
0.000 |
| z |
0.000 |
0.000 |
-46.922 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.197 |
-0.136 |
0.000 |
| y |
-0.136 |
0.248 |
0.000 |
| z |
0.000 |
0.000 |
-0.446 |
|
| Polar |
|---|
| 3z2-r2 | -0.892 |
|---|
| x2-y2 | -0.034 |
|---|
| xy | -0.136 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.695 |
0.450 |
0.000 |
| y |
0.450 |
12.163 |
0.000 |
| z |
0.000 |
0.000 |
12.611 |
<r2> (average value of r
2) Å
2
| <r2> |
224.665 |
| (<r2>)1/2 |
14.989 |