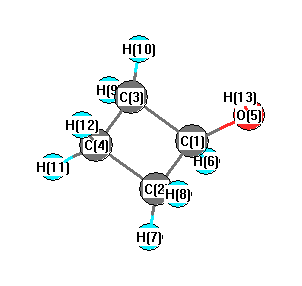
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3863 |
3697 |
19.24 |
|
|
|
| 2 |
A |
3150 |
3015 |
54.00 |
|
|
|
| 3 |
A |
3132 |
2998 |
21.06 |
|
|
|
| 4 |
A |
3092 |
2959 |
21.44 |
|
|
|
| 5 |
A |
3090 |
2958 |
27.22 |
|
|
|
| 6 |
A |
3065 |
2934 |
16.16 |
|
|
|
| 7 |
A |
1511 |
1446 |
9.49 |
|
|
|
| 8 |
A |
1485 |
1422 |
5.88 |
|
|
|
| 9 |
A |
1431 |
1369 |
71.71 |
|
|
|
| 10 |
A |
1334 |
1277 |
0.88 |
|
|
|
| 11 |
A |
1272 |
1217 |
27.10 |
|
|
|
| 12 |
A |
1216 |
1164 |
0.28 |
|
|
|
| 13 |
A |
1165 |
1115 |
115.09 |
|
|
|
| 14 |
A |
1085 |
1039 |
48.72 |
|
|
|
| 15 |
A |
1001 |
958 |
12.16 |
|
|
|
| 16 |
A |
918 |
879 |
1.30 |
|
|
|
| 17 |
A |
760 |
727 |
3.52 |
|
|
|
| 18 |
A |
601 |
575 |
2.68 |
|
|
|
| 19 |
A |
452 |
432 |
4.31 |
|
|
|
| 20 |
A |
187 |
179 |
1.70 |
|
|
|
| 21 |
A |
3132 |
2998 |
40.63 |
|
|
|
| 22 |
A |
3062 |
2931 |
37.38 |
|
|
|
| 23 |
A |
1470 |
1407 |
2.38 |
|
|
|
| 24 |
A |
1296 |
1240 |
0.10 |
|
|
|
| 25 |
A |
1255 |
1201 |
0.22 |
|
|
|
| 26 |
A |
1245 |
1191 |
0.48 |
|
|
|
| 27 |
A |
1188 |
1137 |
0.01 |
|
|
|
| 28 |
A |
1058 |
1012 |
8.85 |
|
|
|
| 29 |
A |
947 |
907 |
4.79 |
|
|
|
| 30 |
A |
935 |
895 |
0.16 |
|
|
|
| 31 |
A |
787 |
753 |
0.62 |
|
|
|
| 32 |
A |
401 |
384 |
37.66 |
|
|
|
| 33 |
A |
290 |
278 |
68.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25436.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24345.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.057 |
|
|
|
| 2 |
C |
-0.239 |
|
|
|
| 3 |
C |
-0.239 |
|
|
|
| 4 |
C |
-0.220 |
|
|
|
| 5 |
O |
-0.345 |
|
|
|
| 6 |
H |
0.124 |
|
|
|
| 7 |
H |
0.110 |
|
|
|
| 8 |
H |
0.105 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
H |
0.105 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.113 |
|
|
|
| 13 |
H |
0.204 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.281 |
-1.518 |
0.000 |
1.544 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.774 |
1.213 |
0.000 |
| y |
1.213 |
-36.202 |
0.000 |
| z |
0.000 |
0.000 |
-32.171 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.413 |
1.213 |
0.000 |
| y |
1.213 |
-6.229 |
0.000 |
| z |
0.000 |
0.000 |
-0.183 |
|
| Polar |
|---|
| 3z2-r2 | -0.367 |
|---|
| x2-y2 | 8.428 |
|---|
| xy | 1.213 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.199 |
-0.443 |
0.000 |
| y |
-0.443 |
7.447 |
0.000 |
| z |
0.000 |
0.000 |
7.376 |
<r2> (average value of r
2) Å
2
| <r2> |
114.322 |
| (<r2>)1/2 |
10.692 |