return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6 (1-Methylcyclopropene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-155.645941
Energy at 298.15K 
HF Energy-154.922923
Nuclear repulsion energy108.755021
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3331 3162 0.72 116.15 0.30 0.46
2 A' 3175 3014 11.09 74.28 0.72 0.84
3 A' 3112 2955 60.59 144.35 0.13 0.23
4 A' 3086 2930 18.27 198.80 0.01 0.02
5 A' 1854 1760 10.57 32.82 0.16 0.28
6 A' 1547 1469 4.78 11.02 0.36 0.53
7 A' 1509 1432 9.23 11.16 0.62 0.77
8 A' 1413 1341 0.21 4.26 0.62 0.77
9 A' 1213 1152 3.19 7.12 0.37 0.54
10 A' 1099 1043 1.99 3.64 0.12 0.21
11 A' 1072 1018 33.66 1.35 0.29 0.45
12 A' 999 949 6.16 7.78 0.26 0.41
13 A' 955 907 14.44 4.63 0.43 0.60
14 A' 689 654 0.57 13.18 0.59 0.74
15 A' 325 308 1.66 1.50 0.75 0.85
16 A" 3176 3016 35.28 88.64 0.75 0.86
17 A" 3150 2991 10.26 72.44 0.75 0.86
18 A" 1506 1429 8.32 7.34 0.75 0.86
19 A" 1135 1078 1.26 1.26 0.75 0.86
20 A" 1059 1006 0.12 2.39 0.75 0.86
21 A" 1005 955 2.54 3.17 0.75 0.86
22 A" 733 695 29.38 0.54 0.75 0.86
23 A" 293 278 11.90 0.53 0.75 0.86
24 A" 165 157 0.34 0.29 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 18799.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17848.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.69591 0.21430 0.17475

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.179 0.000
C2 -0.047 -1.321 0.000
C3 1.152 -0.412 0.000
C4 -0.837 1.384 0.000
H5 -0.316 -1.843 0.909
H6 -0.316 -1.843 -0.909
H7 2.220 -0.368 0.000
H8 -0.233 2.285 0.000
H9 -1.482 1.393 -0.875
H10 -1.482 1.393 0.875

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.50141.29471.46722.24012.24012.28632.11832.10602.1060
C21.50141.50472.81881.08241.08242.45883.61103.19293.1929
C31.29471.50472.67992.24272.24271.06883.03123.31073.3107
C41.46722.81882.67993.39373.39373.52361.08461.08681.0868
H52.24011.08242.24273.39371.81863.07104.22803.87553.4406
H62.24011.08242.24273.39371.81863.07104.22803.44063.8755
H72.28632.45881.06883.52363.07103.07103.61294.19184.1918
H82.11833.61103.03121.08464.22804.22803.61291.76661.7666
H92.10603.19293.31071.08683.87553.44064.19181.76661.7493
H102.10603.19293.31071.08683.44063.87554.19181.76661.7493

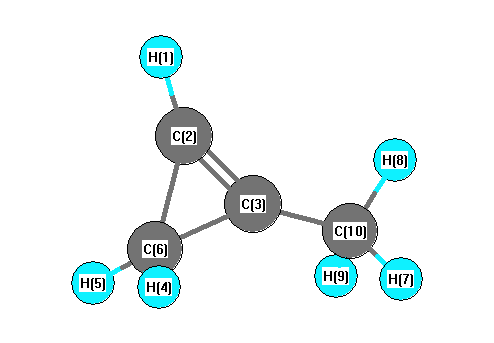
picture of 1-Methylcyclopropene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 51.022 H1 C2 C6 119.327
C2 C3 C6 24.891 C2 C3 C10 72.292
C2 C6 C3 35.809 C2 C6 H4 49.787
C2 C6 H5 32.853 C3 C2 C6 119.300
C3 C6 H4 52.033 C3 C6 H5 66.081
C3 C10 H7 9.315 C3 C10 H8 65.271
C3 C10 H9 74.681 H4 C6 H5 74.458
C6 C3 C10 86.250 H7 C10 H8 58.948
H7 C10 H9 77.956 H8 C10 H9 60.322
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability