Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -959.189353 |
| Energy at 298.15K | |
| HF Energy | -959.189353 |
| Nuclear repulsion energy | 126.653663 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3283 |
3142 |
0.10 |
|
|
|
| 2 |
A1 |
771 |
738 |
8.11 |
|
|
|
| 3 |
A1 |
311 |
298 |
0.38 |
|
|
|
| 4 |
B1 |
325i |
311i |
41.51 |
|
|
|
| 5 |
B2 |
1235 |
1182 |
50.42 |
|
|
|
| 6 |
B2 |
944 |
903 |
150.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3109.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 2976.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.666 |
| H2 |
0.000 |
0.000 |
1.739 |
| Cl3 |
0.000 |
1.465 |
-0.169 |
| Cl4 |
0.000 |
-1.465 |
-0.169 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0731 | 1.6863 | 1.6863 |
H2 | 1.0731 | | 2.4057 | 2.4057 | Cl3 | 1.6863 | 2.4057 | | 2.9299 | Cl4 | 1.6863 | 2.4057 | 2.9299 | |
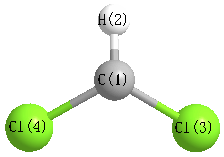 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.687 |
|
Cl3 |
C1 |
Cl4 |
120.627 |
| Cl4 |
C1 |
H2 |
119.687 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.133 |
|
|
|
| 2 |
H |
0.174 |
|
|
|
| 3 |
Cl |
-0.020 |
|
|
|
| 4 |
Cl |
-0.020 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.877 |
0.877 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.568 |
0.000 |
0.000 |
| y |
0.000 |
-30.635 |
0.000 |
| z |
0.000 |
0.000 |
-28.732 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.885 |
0.000 |
0.000 |
| y |
0.000 |
-0.485 |
0.000 |
| z |
0.000 |
0.000 |
2.370 |
|
| Polar |
|---|
| 3z2-r2 | 4.739 |
|---|
| x2-y2 | -0.933 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.282 |
0.000 |
0.000 |
| y |
0.000 |
7.375 |
0.000 |
| z |
0.000 |
0.000 |
4.392 |
<r2> (average value of r
2) Å
2
| <r2> |
98.556 |
| (<r2>)1/2 |
9.928 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -959.189865 |
| Energy at 298.15K | -959.190634 |
| HF Energy | -959.189865 |
| Nuclear repulsion energy | 126.501574 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3247 |
3108 |
0.81 |
|
|
|
| 2 |
A' |
775 |
742 |
12.13 |
|
|
|
| 3 |
A' |
437 |
418 |
26.35 |
|
|
|
| 4 |
A' |
303 |
290 |
1.15 |
|
|
|
| 5 |
A" |
1239 |
1185 |
41.94 |
|
|
|
| 6 |
A" |
918 |
879 |
169.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3459.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 3311.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.686 |
0.000 |
| H2 |
-0.438 |
1.664 |
0.000 |
| Cl3 |
0.011 |
-0.170 |
1.462 |
| Cl4 |
0.011 |
-0.170 |
-1.462 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0759 | 1.6943 | 1.6943 |
H2 | 1.0759 | | 2.3881 | 2.3881 | Cl3 | 1.6943 | 2.3881 | | 2.9240 | Cl4 | 1.6943 | 2.3881 | 2.9240 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
117.345 |
|
Cl3 |
C1 |
Cl4 |
119.279 |
| Cl4 |
C1 |
H2 |
117.345 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.123 |
|
|
|
| 2 |
H |
0.168 |
|
|
|
| 3 |
Cl |
-0.023 |
|
|
|
| 4 |
Cl |
-0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.420 |
0.803 |
0.000 |
0.906 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.348 |
-0.802 |
0.000 |
| y |
-0.802 |
-29.075 |
0.000 |
| z |
0.000 |
0.000 |
-30.723 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.450 |
-0.802 |
0.000 |
| y |
-0.802 |
1.961 |
0.000 |
| z |
0.000 |
0.000 |
-0.511 |
|
| Polar |
|---|
| 3z2-r2 | -1.022 |
|---|
| x2-y2 | -2.274 |
|---|
| xy | -0.802 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.338 |
-0.116 |
0.000 |
| y |
-0.116 |
4.420 |
0.000 |
| z |
0.000 |
0.000 |
7.424 |
<r2> (average value of r
2) Å
2
| <r2> |
98.423 |
| (<r2>)1/2 |
9.921 |
